Home » Blogs from Patient Guard
Blogs
Here we provide you with key information related to the Medical Device and In Vitro Diagnostic (IVD) industry through Patient Guard’s blogs. We provide invaluable content that delves into the regulation and Quality Assurance of Medical Device and IVDs, with focus on navigating the regulatory landscapes of the United Kingdom (UK), the European Union (EU), and the United States (US).
At Patient Guard, we are not just a source of information; we are your ultimate partner in understanding and maneuvering through the complexities of Medical Device and IVD regulatory requirements, as well as Quality Assurance. Our blogs serve as a beacon, illuminating the latest updates and insights crucial for professionals and organizations operating in these highly regulated sectors.
Patient Guard is not just a repository of information; we are your ally in navigating the ever-evolving landscape of Medical Device and IVD regulations. Join us on this educational journey, where compliance meets excellence.
Categories
FDA's Quality Management System Regulation (QMSR) is now in effect. This guide explains how QMSR incorporates ISO 13485:2016, what FDA-specific requirements remain, how FDA inspections have changed and what medical device manufacturers need to do to maintain US compliance....
The MHRA has confirmed plans to recognise CE-marked medical devices indefinitely in Great Britain, removing the planned 2030 deadline for qualifying EU MDR and IVDR devices. Discover what this significant regulatory announcement means for manufacturers, UKCA marking, UK Responsible Person requirements and future access to the UK medical device market....
Understand FDA medical device classification and the differences between Class I, Class II and Class III devices. Learn how to identify your FDA product code, regulatory controls, 510(k) exemptions and the appropriate pathway to the US market....
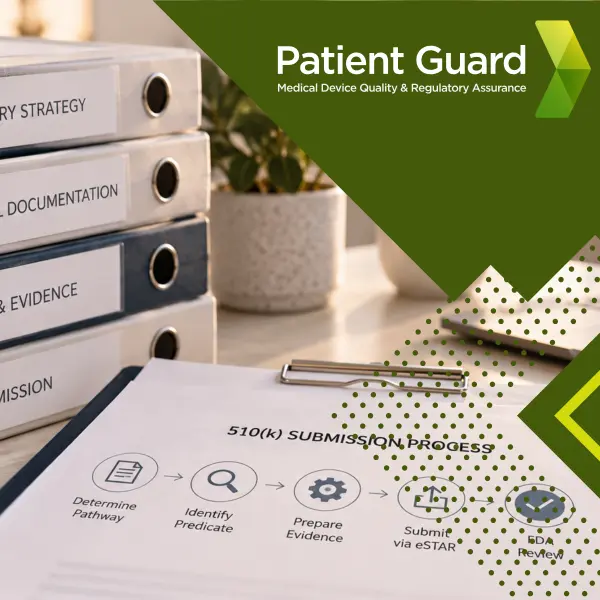
Does your medical device require FDA 510(k) clearance? This step-by-step guide explains the 510(k) process, substantial equivalence, predicate devices, testing, eSTAR submissions, FDA review and what manufacturers need to do after clearance....
Planning to place a medical device on the US market? This complete guide explains FDA medical device regulations, including device classification, 510(k), De Novo and PMA pathways, QMSR and ISO 13485, establishment registration, device listing, U.S. Agent requirements, UDI and post-market obligations....
Preparing to sell digital health technology to the NHS requires more than completing DTAC. This guide explains how manufacturers can build a procurement-ready position by aligning regulatory compliance, clinical safety, data protection, cybersecurity, interoperability, accessibility and commercial evidence for NHS market access.
...