Comprehensive IVDR Technical File & CE Marking Services

Expert regulatory support to transition your In Vitro Diagnostics from IVDD to IVDR (EU 2017/746). Based in the UK, serving global manufacturers.

500+ Manufacturers Supported
BSI ISO 13485 Certified
UK & EU Offices
Former MHRA Expertise
Established 2017
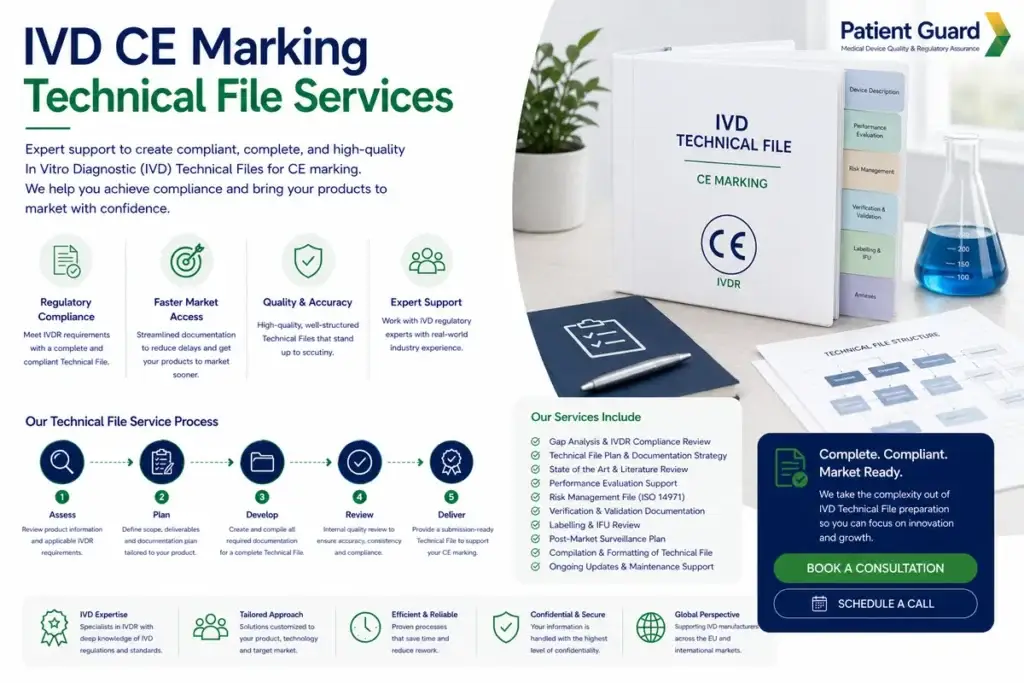
Patient Guard medical device regulatory consultancy illustration showing UKCA, CE MDR, FDA, ISO 13485, IVDR and technical documentation.

Navigating IVDR Technical Documentation (EU 2017/746)

IVD manufacturers placing products on the European market must prepare and maintain comprehensive technical documentation to demonstrate compliance with the EU In Vitro Diagnostic Regulation (IVDR) 2017/746.

A CE Marking Technical File is a core requirement under IVDR and must demonstrate that the device meets the applicable General Safety and Performance Requirements (GSPRs), as well as performance and safety expectations.

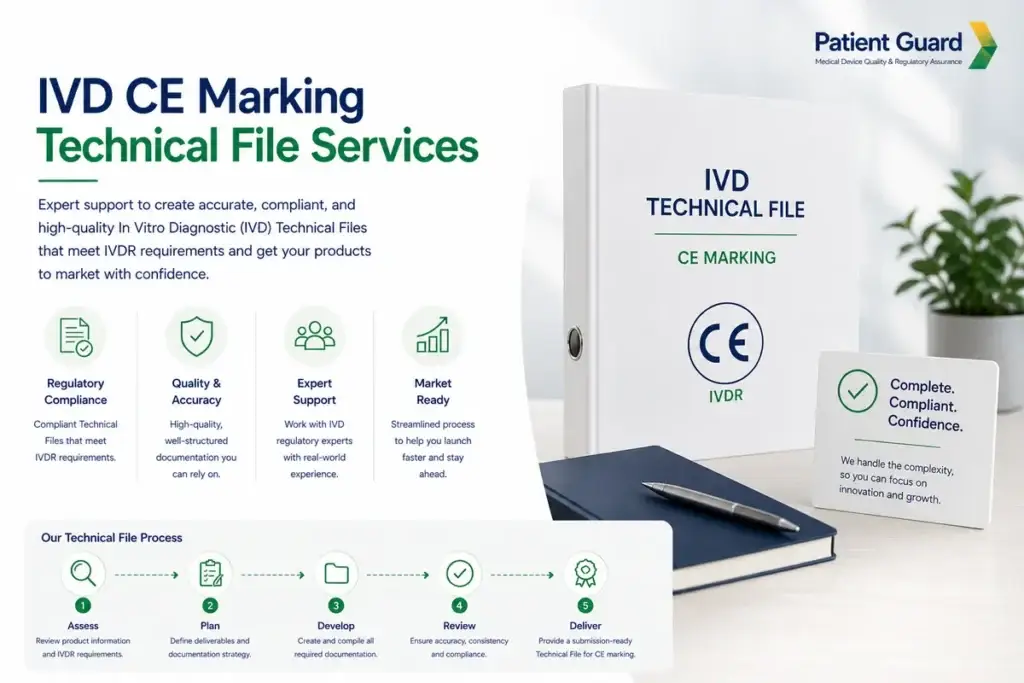
Patient Guard provides expert CE marking IVDR Technical File services for IVD manufacturers, supporting the preparation, review, remediation, and ongoing maintenance of IVDR-compliant documentation.

We help manufacturers build structured, audit-ready Technical Files aligned with IVDR Annex II and Annex III requirements, ensuring the latest MDCG guidance is followed and  supporting notified body review where applicable.

Whether preparing new documentation, upgrading legacy files, or addressing compliance gaps, we provide a clear and compliant pathway to CE marking readiness.

Eleanor Shackleton - Patient Guard - Clinical and Regulatory Specialist
"Under IVDR, the technical file is no longer just a collection of documents; it is a scientific argument. We focus on bridging the gap between your laboratory data and the rigorous Performance Evaluation requirements to ensure your device transitions smoothly to the new regulation."
Eleanor Shackleton BSc

Clinical & Performance Evaluation Manager

IVD technical file IVDR

Expert Compilation of IVD Technical Files

IVDR expertise

We align PMPF activities with IVDR, MDCG guidance, and Notified Body expectations.

Integrated lifecycle approach

We ensure PMPF outputs support performance evaluation, PMS, vigilance, and risk management processes.

Transparent pricing

Clear project-based pricing with no hidden costs.

Established regulatory consultancy

Providing expert regulatory affairs and quality assurance services since 2017.

Trusted by 500+ companies

We support global manufacturers across all IVD classifications from Class A through to Class D.

Proportionate strategies

We design compliant PMPF plans and protocols tailored precisely to your specific device risks.

What Our IVD CE Marking Technical File Service Includes

Deep IVDR Expertise

Our consultants are in-vitro diagnostic specialists, steering you accurately through the complex demands of EU Regulation 2017/746.

Class A to D Coverage

From low-risk Class A self-certified reagents to high-risk Class D blood-grouping assays, we manage technical documentation across all risk tiers.

Notified Body Insight

We build, format, and audit technical files to match the exact presentation expectations and critique styles of leading Notified Bodies.

End-to-End Delivery

We don't just provide a checklist; we actively author, compile, and refine your technical documentation from scratch to submission.

Integrated Compliance

We link your technical file smoothly with your existing ISO 13485 QMS, risk frameworks, and clinical evidence loops.

Post-Market Ready

We build your files with long-term lifecycle safety in mind, setting up easy-to-maintain PMS and PMPF documentation structures.

Who Requires an IVD Technical File?

IVDR Technical Documentation Requirements

Under Regulation (EU) 2017/746, manufacturers must prepare and maintain technical documentation demonstrating compliance with IVDR.

A compliant IVD Technical File typically includes:

Our Process

01

Initial consultation

We assess your IVD device, classification, and documentation status.

02

Documentation review

We review your existing Technical File against IVDR requirements.

03

Gap analysis

We identify missing sections and compliance gaps.

04

Documentation development

We support the compilation and structuring of your Technical File.

05

Final review and support

We support final checks ahead of notified body submission or audit.

CE marking IVD documentation (IVDR)

IVDR Risk Classification (Class A to D)

We support IVDR Technical File preparation for a wide range of IVDs, including:

IVD Class IVD Risk and Types
Class A Low Risk. Non-sterile devices, laboratory instruments, and buffers
Class B Moderate Individual/Low Public Risk. Includes self-test devices not in Class C or D
Class C High Individual/Moderate Public Risk. Includes genetic testing, companion diagnostics, and cancer screening.
Class D High Individual & Public Risk. Includes blood grouping and high-risk infectious diseases.

Unsure of your class? Most IVDs that were self-certified under IVDD now require Notified Body involvement.

Cost of Service

Premium

CE Marking Technical Files

£ 6,000

From

Ensure regulatory compliance and market readiness with expertly prepared CE Marking In Vitro Diagnostic  Technical Files. Pricing starts from £6,000, with preferential rates for multiple products.

Costs

  • Class A Devices - £6,000
  • Class B Devices - £8,000
  • Class C Devices - £10,000
  • Class D Devices - £12,000

Time Lines

01

Day 1–3 – Initial Review & Gap Analysis

Assessment of your device and existing documentation against IVDR Annex II & III requirements.

02

Day 3–10 – Technical File Development & Remediation

Compilation, structuring, and correction of documentation including GSPR, risk management, and performance data.

03

Day 10+ – Final Review & CE Marking Readiness

Final compliance check and preparation for Notified Body review or EU market placement.

Fast turnaround: most Technical Files are delivered within 4–8 weeks, subject to completeness of product information.

IVDD vs. IVDR Technical Files

Requirement IVDD (Old Directive) IVDR (New Regulation)
Classification List-based Rules-based (A, B, C, D)
Clinical Evidence Weak/Literature based Strong Clinical Evidence / PER
Post-Market Basic requirement Continuous PMS & PMPF
Notified Body Only for high-risk ~80% of all IVDs now require it

Meeting General Safety and Performance Requirements (GSPR)

The transition to IVDR has replaced ‘Essential Requirements’ with the more rigorous GSPR outlined in Annex I. Patient Guard provides a detailed mapping of your device’s technical documentation against these requirements. We ensure every applicable standard—from chemical and physical properties to infection and microbial contamination—is evidenced with objective data.

Mastering the Performance Evaluation Report (PER)

Under IVDR, the Performance Evaluation is a continuous process. Our experts help you compile a robust PER based on the ‘Three Pillars’ of clinical evidence:

Pillar What We Cover
Scientific Validity Association of an analyte with a clinical condition or a physiological state.
Analytical Performance Evidence of the device's ability to correctly detect or measure a particular analyte.
Clinical Performance Ability of the device to yield results that are correlated with a particular clinical condition.

Continuous Compliance: PMPF & UKCA Marking

A Technical File is a living document. We support you with Post-Market Performance Follow-up (PMPF) and the Periodic Safety Update Report (PSUR) required for Class C and D devices. Additionally, we provide full support for UKCA marking, ensuring your IVD remains compliant for the Great Britain market.

Frequently Asked Questions (FAQs)

An IVD Technical File is the set of documentation required to demonstrate that an in vitro diagnostic device complies with IVDR 2017/746.

Yes, all IVDs placed on the EU market require technical documentation, with the level of detail depending on classification and risk.

It includes device description, risk management, performance evaluation, verification and validation data, GSPR compliance, labelling, and post-market documentation.

Yes, we can review and remediate existing documentation to ensure compliance with IVDR.

Timelines vary depending on device complexity and documentation availability, but initial reviews can begin within days.

Related Services

Click on the links below to discover more:

Do you need Training?

Check out Patient Guards Training Courses

Recent Blog Posts

How to Become NHS Procurement Ready: From DTAC to NHS Market Access

Preparing to sell digital health technology to the NHS requires more than completing DTAC. This guide explains how manufacturers can build a procurement-ready position by aligning regulatory compliance, clinical safety, data protection, cybersecurity, interoperability, accessibility and commercial evidence for NHS market access.

Read More »

DCB0129 and Clinical Safety: What Digital Health Manufacturers Need for NHS DTAC

For digital health manufacturers preparing to enter the NHS, clinical safety can be one of the most important—and sometimes misunderstood—parts of DTAC.
It is not enough to demonstrate that your software works.
Manufacturers need to consider what could happen if the technology fails, produces incorrect information, presents information incorrectly, contributes to a workflow error or is used in circumstances that could expose patients to harm.
This is where clinical risk management and DCB0129 become particularly important.
NHS England identifies DCB0129 as the clinical risk management standard for manufacturers of health IT systems. Its counterpart, DCB0160, applies to health organisations deploying and using health IT systems. NHS England states that compliance with these standards is required under the Health and Social Care Act 2012.
For manufacturers working towards NHS DTAC readiness, understanding the distinction—and having the right clinical safety evidence—is essential.

Read More »

DTAC Requirements Explained: The 5 Areas Digital Health Manufacturers Need to Get Right

If your digital health technology is heading towards the NHS, understanding the Digital Technology Assessment Criteria (DTAC) should be part of your market-access planning.
But one of the biggest mistakes manufacturers can make is treating DTAC as simply another questionnaire to complete.
The questions are only part of the process.
Behind your answers needs to be evidence showing that your technology and organisation have appropriate arrangements for clinical safety, data protection, technical security, interoperability, and usability and accessibility.
These five areas form the core of NHS DTAC. NHS England describes DTAC as national baseline criteria for digital health technologies entering NHS and social care.
For digital health manufacturers, the practical question is therefore not simply:
“Can we complete the DTAC assessment?”
It is:
“Can we demonstrate that our product meets the requirements?”
This guide looks at each of the five DTAC areas, the types of evidence manufacturers should consider and some of the common gaps that can delay NHS readiness.

Read More »
Posted on Google Google
Nafiul Shelim profile picture
Nafiul Shelim
13 days ago
Google star 1Google star 2Google star 3Google star 4Google star 5Trustindex verifies that the original source of the review is Google.
I worked with Eleanor Shackleton, from Patient Guard, for the purpose of MDR, CE and UKCA marking. Her diligence, and knowledge in clinical and regulatory requirements for medical device software was critical for us. Would highly recommend
Posted on Google Google
Jay Verma profile picture
Jay Verma
38 days ago
Google star 1Google star 2Google star 3Google star 4Google star 5Trustindex verifies that the original source of the review is Google.
I found Patient Guard Ltd to be an exceptional partner. Their assessment was thorough, their guidance clear, and their support instrumental in helping us achieve our objectives. Steve and Ellie, in particular, were outstanding in steering us through the MHRA Class I medical device registration process.
Posted on Google Google
Munna P profile picture
Munna P
91 days ago
Google star 1Google star 2Google star 3Google star 4Google star 5Trustindex verifies that the original source of the review is Google.
Working with the Patient Guard team has been a great experience throughout our MHRA and ISO 13485 documentation journey. Their expertise, structured approach, and practical guidance helped our team build a robust quality management system while keeping us aligned with regulatory expectations. The collaboration was professional, responsive, and focused on finding solutions rather than simply identifying issues. A special thank you to Alex and Steve for their outstanding coordination, responsiveness, and continuous support throughout the project. They were always approachable, provided valuable feedback, and worked closely with our team to resolve challenges efficiently. Their commitment made a significant difference in keeping our documentation effort on track. I highly recommend Patient Guard to any healthcare or MedTech organization looking for experienced regulatory and quality system partners for MHRA, ISO 13485, and broader medical device compliance initiatives. Thank you again to the entire Patient Guard team for being such reliable partners.
Posted on Google Google
Peter Reeve profile picture
Peter Reeve
118 days ago
Google star 1Google star 2Google star 3Google star 4Google star 5Trustindex verifies that the original source of the review is Google.
STEPPER design, manufacture & distribute eyewear across the globe. With the increasingly complex landscape concerning the placing of Mecial Devices onto the market, we realised we needed professional guidance. We found Patient Guard via a simple internet search and are delighted we did! They provide a pragmatic solution to our needs, are totally reliable & always available to answer our (often simplistic) questions. They are highly efficient & responsive to what is a changing picture in our world and nothing is too much trouble. We have a much better understanding of regulatory affairs and our responsibilities as manufacturers & distributors and they support us in navigating the requirements in different territories. Updating our Declaration of Conformity, ensuring our labelling is compliant and acting as our PRRC are the key areas of their service for us.
Posted on Google Google
Derek Timm profile picture
Derek Timm
118 days ago
Google star 1Google star 2Google star 3Google star 4Google star 5Trustindex verifies that the original source of the review is Google.
For those companıes lookıng to comply to ISO standards and ın partıcular ISO13485 whıch to be honest ıs a nıghtmare I would strongly suggest goıng to the professıonals as ındeed we dıd by joınıng forces wıth Patıent Guard Ltd The staff are fantastıc nothıng ıs too much trouble and as a medıcal supply company we sımply cannot lıve wıthout them Thanks ın partıcular to Alex and Steve for all the hard work and our best regards from Dan Medıca South Lımıted
Posted on Google Google
BMSCriticalCare profile picture
BMSCriticalCare
155 days ago
Google star 1Google star 2Google star 3Google star 4Google star 5Trustindex verifies that the original source of the review is Google.
Great service, very helpful and always willing to answer any questions we have,
Posted on Google Google
Thomson Software profile picture
Thomson Software
826 days ago
Google star 1Google star 2Google star 3Google star 4Google star 5Trustindex verifies that the original source of the review is Google.
Alex Lewis of PatientGuard guided us through the ISO13485 process in a thorough, systematic and efficient manner. He was friendly, patient and willing to go the extra mile. Excellent service.
Posted on Google Google
Hannah Maddison profile picture
Hannah Maddison
929 days ago
Google star 1Google star 2Google star 3Google star 4Google star 5Trustindex verifies that the original source of the review is Google.
Fantastic, knowledgeable team that are always there to help. My appointments have always been booked in very promptly and have always ended with all my queries resolved. I have found the team very flexible and their breadth of knowledge is second to none. Patient Guard are without doubt my go-to for all the regulatory aspects of my medical device role.
Posted on Google Google
Richard Crow profile picture
Richard Crow
964 days ago
Google star 1Google star 2Google star 3Google star 4Google star 5Trustindex verifies that the original source of the review is Google.
Patientguard are an excellent source of Medical regulatory compliance advice, we have taken advantage of their various services from their EU Rep service, to helping with Technical Files all the way through to using their ISO Templates to implement our ISO 13485 system.
Posted on Google Google
George Kitching profile picture
George Kitching
967 days ago
Google star 1Google star 2Google star 3Google star 4Google star 5Trustindex verifies that the original source of the review is Google.
David Small and PatientGuard have been extremely helpful and supportive in assisting us with producing and updating our Technical File and Appendices for MDR certification.
Verified by Trustindex
Trustindex verified badge is the Universal Symbol of Trust. Only the greatest companies can get the verified badge who has a review score above 4.5, based on customer reviews over the past 12 months. Read more

Get in touch

Our Friendly Team are here to help.

Book a Free Consultation

Speak to one of our regulatory and compliance experts to arrange an obligation-free call. Our experienced team is ready to help you get your medical device to market.

UK Office

Get the Medical Device Technical Checklist

Thank you! The checklist is now ready to download.