Updated: 7th October 2026
Reviewed by: David Small BSc (Hons), MSc, MTOPRA (Founder and CEO)
Medical Device Consultancy for UK and EU Regulatory Compliance
Medical device manufacturers operate in one of the most highly regulated sectors in the world. A successful product launch depends not only on innovation, but also on demonstrating compliance with applicable UK and EU medical device regulations, quality management system requirements, and safety and performance expectations.
For many organisations, navigating these requirements creates a significant challenge. Internal teams may have strong technical expertise but limited regulatory capacity. Start-ups may need help understanding their first route to market, while established manufacturers may require specialist support with Technical Documentation, ISO 13485, clinical evaluation, biological evaluation, risk management, Notified Body requirements, CE marking, UKCA marking, post-market surveillance or regulatory strategy.
A specialist medical device consultancy provides access to experienced regulatory and quality professionals who can help manufacturers understand the requirements that apply to their devices, identify compliance gaps, develop regulatory strategies, prepare documentation and manage regulatory activities in a structured and practical way.
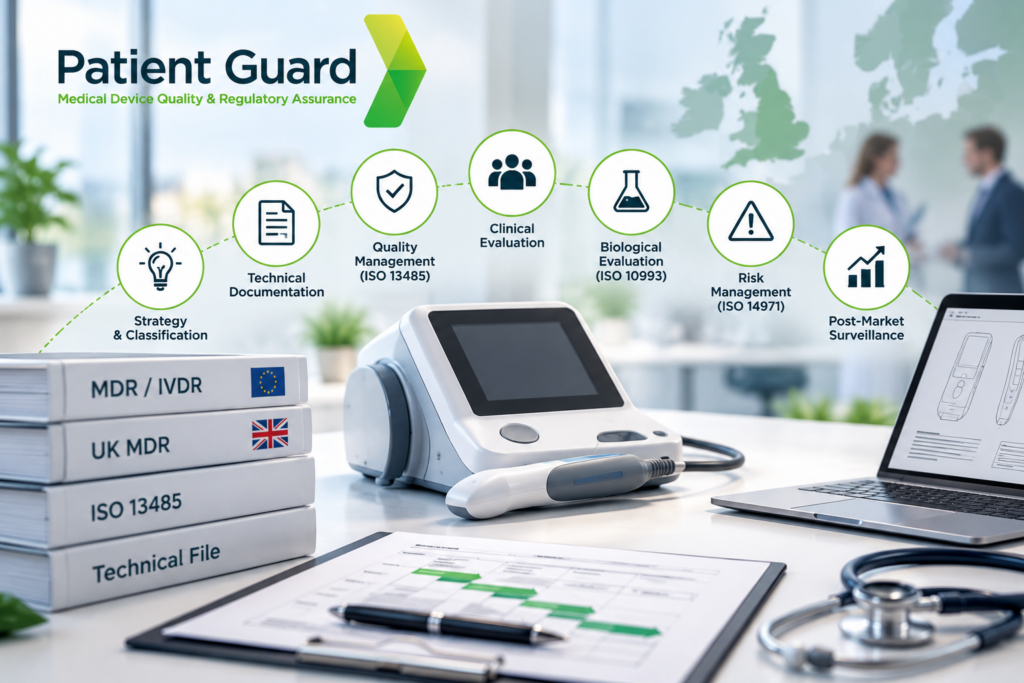
At Patient Guard, our medical device consultants support manufacturers throughout the product lifecycle. This can begin with early-stage device classification and regulatory planning and continue through quality management, technical documentation and clinical evidence to market access, certification, post-market surveillance and ongoing regulatory compliance in the UK and EU.
Need Help Understanding the EU MDR?
Our medical device consultancy services support manufacturers throughout every stage of EU MDR compliance. If you're looking for a complete overview of Regulation (EU) 2017/745, including classification, Technical Documentation, Clinical Evaluation, UDI, EUDAMED and CE marking, read our Complete Guide to the EU MDR.
Manufacturing In Vitro Diagnostic Devices?
Patient Guard supports IVD manufacturers with every aspect of IVDR compliance. Learn about classification, Performance Evaluation, Technical Documentation, Notified Bodies, UDI and post-market requirements in our Complete Guide to the EU IVDR.
Need Guidance on UK Medical Device Regulations?
Whether you're placing medical devices on the Great Britain market or preparing for future UK regulatory changes, our consultants can help. Learn about UKCA marking, MHRA requirements, UK Responsible Persons and UK device registration in our Complete Guide to UK Medical Device Regulations.
What Is Medical Device Consultancy?
Medical device consultancy is the provision of specialist regulatory, quality assurance and technical expertise to organisations developing, manufacturing or marketing medical devices and in vitro diagnostic (IVD) devices.
Rather than maintaining a large in-house regulatory and quality team, many organisations choose to work with an experienced medical device consultant or consultancy that can provide specialist support when and where it is required.
Medical device consultants can help manufacturers understand complex regulatory requirements, determine appropriate routes to market, develop compliance strategies, prepare technical documentation and navigate interactions with regulatory authorities and Notified Bodies. They may also provide specialist support in areas such as quality management, clinical evaluation, biological evaluation, risk management and post-market surveillance.
The role of a medical device consultancy is ultimately to help manufacturers reduce regulatory risk, avoid unnecessary delays and establish a clear route towards market access and ongoing compliance.
Whether you are developing your first medical device or managing an established international product portfolio, regulatory consultancy can provide additional expertise and capacity throughout the device lifecycle.
What Does a Medical Device Consultant Do?
Medical device consultants provide support across a wide range of regulatory and quality activities, including:
• Developing regulatory strategies for new products.
• Determining whether a product qualifies as a medical device or IVD.
• Medical device and IVD classification.
• Preparing Technical Documentation.
• Clinical Evaluation Reports (CERs).
• Biological Evaluation Reports (BERs).
• Risk Management documentation in accordance with ISO 14971.
• Quality Management System implementation and maintenance to ISO 13485.
• Software lifecycle support under IEC 62304.
• Usability Engineering under IEC 62366-1.
• CE marking and UKCA compliance.
• EU MDR and IVDR compliance.
• UK medical device regulatory support.
• EU Authorised Representative and UK Responsible Person services.
• Internal audits and supplier audits.
• Notified Body preparation and audit support.
• Post-Market Surveillance (PMS) and PMCF.
By providing specialist knowledge across these disciplines, consultants allow manufacturers to focus on innovation while ensuring regulatory requirements are addressed efficiently and effectively.
Why Do Manufacturers Use Medical Device Consultants?
Medical device manufacturers must navigate an increasingly demanding regulatory environment. The EU Medical Device Regulation (MDR), In Vitro Diagnostic Regulation (IVDR), evolving UK medical device requirements, increasing expectations for clinical evidence, and requirements affecting medical device software all require specialist regulatory and quality expertise.
For many organisations, maintaining in-house expertise across every regulatory discipline is neither practical nor cost-effective. Requirements can also vary considerably according to the type and classification of the device, target market and stage of the product lifecycle.
Working with an experienced medical device consultancy enables manufacturers to:
- Access specialist regulatory and quality expertise when required.
- Identify and reduce regulatory and compliance risks.
- Avoid delays and costly compliance mistakes.
- Establish appropriate regulatory strategies and routes to market.
- Prepare for Notified Body assessments and certification activities.
- Develop and maintain effective quality management systems.
- Address specialist areas such as clinical evaluation, biological evaluation, risk management and software compliance.
- Support UK, EU and international market access.
- Maintain regulatory compliance throughout the post-market phase.
Medical device consultancy can therefore be used for anything from a single specialist project to ongoing regulatory and quality support. Consultants can also work alongside an existing regulatory team, providing additional capacity or expertise where internal resources are limited.
This flexible approach allows manufacturers to access experienced professionals when needed without necessarily recruiting permanent specialists for every area of medical device regulation and quality assurance.
Medical Device Consultancy Throughout the Product Lifecycle
Medical device consultancy can provide regulatory and quality support at every stage of the medical device lifecycle, from initial concept and development through to market access and ongoing post-market compliance.
During concept development, medical device consultants can help manufacturers define the intended purpose of the device, determine its regulatory classification, identify applicable regulatory requirements and develop an appropriate market access strategy for the UK, EU and other target markets.
During design and development, consultancy support may include risk management, usability engineering, biological evaluation, medical device software compliance and the development or implementation of an appropriate quality management system.
Before market launch, consultants can support the preparation and review of Technical Documentation, Clinical Evaluation Reports, declarations of conformity and other documentation required to demonstrate regulatory compliance. Where applicable, they can also support manufacturers through Notified Body assessment and regulatory submission activities.
Following market access, consultancy support can continue through Post-Market Surveillance, vigilance, change management, regulatory updates, periodic reporting and preparation for Notified Body surveillance audits.
Integrating regulatory expertise throughout the product lifecycle can help manufacturers identify potential compliance issues earlier, maintain consistency between regulatory documentation and product development, and reduce the risk of delays or costly remediation later in the regulatory process.

When Should You Hire a Medical Device Consultancy?
Many manufacturers assume that regulatory consultants are only needed when preparing for CE marking, UKCA marking or a Notified Body assessment. In reality, involving experienced regulatory and quality professionals earlier in the product lifecycle can help identify potential problems before they result in delays, additional testing or costly remediation.
A medical device consultancy can be particularly valuable when an organisation lacks specialist expertise in a particular regulatory area, needs additional capacity for a defined project or is entering a new regulatory market.
Whether you are developing your first medical device or managing an established international product portfolio, external consultants can work alongside your existing team to provide specialist expertise at different stages of the regulatory journey.
Some of the most common situations where manufacturers benefit from medical device consultancy support include:
Developing a New Medical Device
Some of the most important regulatory decisions are made during the earliest stages of medical device development. Decisions concerning the device’s intended purpose, classification, design and target markets can influence the regulatory requirements that apply throughout the rest of the project.
At the concept stage, a medical device consultancy can help determine whether the product qualifies as a medical device or IVD, identify the applicable regulatory framework, establish the appropriate classification and develop a regulatory strategy before significant investment is made in design, testing and documentation.
Addressing these questions early can help manufacturers avoid unnecessary testing, inappropriate development pathways, costly redesigns and regulatory delays later in the project.
Typical consultancy support includes:
- Medical device or IVD product determination.
- Intended purpose and claims review.
- Device classification.
- Identification of applicable regulatory requirements.
- UK, EU and international regulatory strategy.
- Regulatory development planning.
Medical Device Start-ups
Medical device start-ups often begin with strong technical or clinical expertise but may have limited experience navigating medical device regulation. Regulatory requirements can have a significant impact on product development, testing requirements, timelines, costs and the eventual route to market.
Working with a medical device consultancy at an early stage can help founders understand the complete regulatory pathway before making significant development commitments or approaching investors, clinical partners, manufacturers or other commercial stakeholders.
Consultants can also help translate regulatory requirements into a realistic development plan, ensuring that activities such as quality management, risk management, verification and validation, clinical evidence and Technical Documentation develop alongside the product rather than being addressed retrospectively.
Typical consultancy support includes:
- Regulatory roadmaps and gap assessments.
- Medical device classification and regulatory strategy.
- Route-to-market planning.
- Development and regulatory milestone planning.
- Quality Management System implementation.
- Technical Documentation planning.
- Identification of testing and evidence requirements.
- Regulatory input to investor or commercial due diligence.
Early regulatory planning can give start-ups a clearer understanding of the time, evidence and resources required to reach the market, helping regulatory strategy become part of the wider business plan rather than an unexpected barrier later in development.
Preparing for CE Marking or UKCA Marking
Achieving CE marking or UKCA marking requires manufacturers to demonstrate that their medical device meets the applicable regulatory requirements. The precise conformity assessment route depends on factors including the device classification, intended purpose and target market.
This involves considerably more than preparing Technical Documentation. Manufacturers may need to establish an appropriate quality management system, demonstrate effective risk management, generate sufficient clinical and biological safety evidence, complete verification and validation activities and prepare the documentation necessary to demonstrate conformity.
A medical device consultancy can help manufacturers identify the applicable requirements, assess existing evidence and documentation, identify gaps and coordinate the regulatory activities required to progress towards market access.
Typical consultancy support includes:
- Technical Documentation preparation and review.
- Clinical Evaluation.
- Biological Evaluation.
- Risk Management to ISO 14971.
- Quality Management Systems and ISO 13485.
- Verification and validation planning.
- Notified Body preparation and support, where applicable.
- CE marking under the EU MDR or IVDR.
- UKCA regulatory compliance.
- Declarations of Conformity and associated regulatory documentation.
Preparing for a Notified Body Assessment
Many manufacturers seek specialist consultancy support before their first Notified Body assessment, during certification activities or when preparing for subsequent surveillance audits.
An independent medical device consultant can review the manufacturer’s quality and regulatory documentation, identify potential gaps or nonconformities and assess whether the organisation is adequately prepared for the assessment process.
This may include reviewing the Quality Management System, Technical Documentation, clinical evidence, risk management documentation and post-market surveillance processes, as well as conducting internal or mock audits to identify weaknesses before they are raised by the Notified Body.
Where findings have already been identified, consultants can also support manufacturers with root cause analysis, corrective and preventive action (CAPA) and regulatory remediation.
Typical consultancy support includes:
- Regulatory and QMS gap assessments.
- Technical Documentation reviews.
- ISO 13485 internal audits.
- Mock Notified Body audits.
- Audit readiness and staff preparation.
- Review of clinical, risk management and PMS documentation.
- Root cause analysis and CAPA support.
- Remediation of audit findings and nonconformities.
- Ongoing regulatory remediation and compliance support.
Early preparation can help manufacturers identify potential issues before the formal assessment, improve audit readiness and reduce the risk of avoidable findings or delays during the certification process.
Managing Changes to an Existing Medical Device
Even relatively small changes to an existing medical device can affect its regulatory compliance. Manufacturers should therefore assess the potential regulatory impact of a change before implementation rather than assuming that existing evidence and documentation remain sufficient.
Changes to materials, components, suppliers, manufacturing processes, software, sterilisation methods, packaging, intended purpose, indications or product claims may require updates to Technical Documentation and could affect existing verification and validation evidence.
Depending on the nature of the change, manufacturers may also need to review or update risk management documentation, clinical evaluation, biological evaluation, usability engineering, software documentation and post-market surveillance activities. Certain changes may also need to be assessed in relation to existing conformity assessment or certification arrangements.
A medical device consultancy can provide an independent regulatory assessment of the proposed change, identify the documentation and evidence affected and help manufacturers determine the regulatory activities required before implementation.
Typical consultancy support includes:
- Regulatory change impact assessments.
- Technical Documentation updates.
- Risk Management File updates.
- Clinical Evaluation updates.
- Biological Evaluation reviews and updates.
- Verification and validation impact assessments.
- Software and usability documentation reviews.
- Labelling and IFU reviews.
- Quality Management System change control.
- Assessment of potential implications for existing certification or conformity assessment.
A structured change assessment helps manufacturers maintain compliance throughout the device lifecycle and reduces the risk of implementing changes that unintentionally affect the device’s regulatory status or supporting evidence.
Expanding a Medical Device into New Markets
Entering a new international market can introduce additional regulatory requirements, even where a medical device is already legally marketed in another jurisdiction. Manufacturers need to understand the regulatory framework, classification rules, conformity assessment requirements, registration obligations and documentation expected in each target market.
For example, a device already CE marked in the European Union may require additional regulatory activities before being placed on the Great Britain market, while manufacturers entering the EU may need to address the requirements of the EU MDR or IVDR, including the appointment of an EU Authorised Representative where applicable.
A medical device consultancy can assess the manufacturer’s existing regulatory documentation and evidence, identify differences between jurisdictions and develop an appropriate strategy for entering the new market. Existing Technical Documentation, clinical evidence, testing and quality system documentation can also be reviewed to determine what may be leveraged and where additional work is required.
Typical consultancy support includes:
- EU MDR and IVDR compliance.
- UK medical device regulatory compliance.
- Regulatory gap assessments between markets.
- Device classification and route-to-market assessment.
- International regulatory strategy.
- Technical Documentation gap analysis.
- EU Authorised Representative and UK Responsible Person requirements.
- Regulatory registration requirements.
- Market access planning.
A structured market expansion strategy can help manufacturers reuse appropriate existing evidence and documentation while identifying market-specific requirements early, reducing duplication and avoiding unnecessary delays when entering new jurisdictions.
Additional Regulatory and Quality Capacity
Not every organisation needs, or has sufficient workload to justify, permanent in-house specialists across every area of medical device regulation and quality assurance. Even manufacturers with established regulatory and quality teams may require additional capacity during periods of growth, increased workload or specialist projects.
A medical device consultancy can work alongside an existing team, providing additional regulatory and quality resources when required. This may involve supporting a defined project, covering a temporary resource gap or providing specialist expertise that is not available internally.
This flexible approach enables manufacturers to access experienced regulatory and quality professionals without necessarily increasing permanent headcount, while allowing internal teams to maintain responsibility for their wider regulatory and quality activities.
Typical consultancy support includes:
- Temporary regulatory or quality resource support.
- Maternity or other extended leave cover.
- Regulatory project management.
- Technical Documentation preparation and remediation.
- Clinical and biological evaluation support.
- Internal auditing.
- Supplier audits.
- Quality Management System implementation and improvement.
- Post-market surveillance activities.
- Specialist regulatory and quality projects.
Consultancy support can therefore range from a single defined project to longer-term retained regulatory and quality support, depending on the manufacturer’s internal resources and changing requirements.
Long-Term Regulatory and Quality Support
Medical device compliance does not end once a product reaches the market. Manufacturers must continue to monitor device performance, maintain Technical Documentation, manage post-market activities and ensure that their Quality Management System and regulatory documentation remain appropriate throughout the device lifecycle.
Manufacturers must also respond to product changes, emerging risks, complaints, vigilance issues, regulatory developments and new post-market information that could affect the continued safety, performance or compliance of their devices.
For this reason, many organisations choose to work with a medical device consultancy on an ongoing basis. Long-term support can provide access to regulatory and quality expertise as requirements arise, while helping manufacturers maintain their systems and documentation rather than addressing compliance issues retrospectively.
Long-term consultancy support can include:
- Regulatory monitoring and updates.
- Post-Market Surveillance (PMS).
- Post-Market Clinical Follow-up (PMCF).
- Vigilance and complaint management support.
- Technical Documentation maintenance and updates.
- Clinical Evaluation and risk management updates.
- Internal audits.
- Management review support.
- Supplier management and supplier audits.
- Quality Management System maintenance.
- Change control and regulatory impact assessments.
- Corrective and Preventive Action (CAPA).
- Continual improvement activities.
Long-term regulatory and quality support can provide manufacturers with continuity throughout the device lifecycle, helping ensure that regulatory documentation, quality systems and post-market activities continue to develop alongside the device and the regulatory environment.
When to Engage a Medical Device Consultancy
The earlier regulatory expertise is introduced into a medical device project, the greater the opportunity to identify potential compliance issues, reduce regulatory risk and avoid unnecessary development work.
While medical device consultants can support manufacturers with remediation and compliance issues later in development, involving regulatory specialists earlier allows regulatory requirements to be considered alongside product design, testing, clinical evidence, quality management and commercial planning.
Early involvement can help manufacturers establish a clearer regulatory pathway, anticipate evidence and documentation requirements and reduce the risk of costly redesigns or delays later in the development process.
However, consultancy support can be valuable at any stage of the medical device lifecycle. Whether you require specialist expertise for a single project, additional capacity for an existing regulatory team or a long-term regulatory and quality partner, the right support can help establish a more structured and predictable route to market and ongoing compliance.

Who Uses Medical Device Consultants?
Medical device consultancy is not limited to large multinational manufacturers. Organisations of all sizes use specialist regulatory and quality expertise to help navigate complex compliance requirements, support product development and achieve and maintain market access.
From medical device start-ups developing their first product to established manufacturers managing international product portfolios, medical device consultants can provide specialist expertise and additional capacity according to the organisation’s regulatory, quality and commercial needs.
Consultancy support can also be valuable for organisations entering the medical device sector, overseas manufacturers seeking access to the UK or EU markets, and established regulatory or quality teams requiring additional expertise for specific projects.
At Patient Guard, we work with organisations across the medical device and in vitro diagnostic sector, providing flexible regulatory and quality consultancy support that can scale alongside the manufacturer and its product portfolio.
Medical Device Start-ups
Medical device start-ups often combine innovative technology and strong technical or clinical expertise with limited in-house regulatory and quality resources. Understanding the regulatory pathway early is particularly important because regulatory requirements can influence product design, development costs, testing requirements, investment needs and timelines to market.
Patient Guard can work alongside founders and development teams to establish a practical regulatory strategy from the outset. This can include determining whether the product qualifies as a medical device or IVD, establishing its classification, identifying applicable regulatory requirements and planning the quality, technical and clinical activities required to reach the target market.
For start-ups seeking investment, a clearly defined regulatory pathway can also help demonstrate that the resources, evidence, costs and development milestones required for market access have been properly considered.
Typical consultancy support includes:
- Medical device or IVD product determination.
- Device classification.
- Regulatory strategy and roadmaps.
- UK and EU route-to-market planning.
- Development and regulatory milestone planning.
- Quality Management System implementation.
- Technical Documentation planning and preparation.
- Identification of testing and evidence requirements.
- Investor and regulatory due diligence support.
Small and Medium-Sized Manufacturers
Small and medium-sized medical device manufacturers often have established engineering, quality or regulatory teams but may not have sufficient internal resources or specialist expertise to manage every regulatory requirement.
As product portfolios grow, new markets are entered or regulatory workloads increase, a medical device consultancy can provide additional capacity and specialist expertise without requiring manufacturers to recruit permanent staff for every regulatory discipline.
Consultants can work alongside existing teams, taking responsibility for defined projects or providing expertise in areas such as clinical evaluation, biological evaluation, Technical Documentation, quality management and regulatory compliance.
This can be particularly valuable when preparing for certification activities, responding to regulatory findings, managing periods of increased workload or completing specialist projects that fall outside the organisation’s normal regulatory activities.
Typical consultancy support includes:
- Technical Documentation preparation and review.
- Clinical Evaluation.
- Biological Evaluation.
- ISO 13485 implementation and QMS support.
- Risk management.
- Internal and supplier audits.
- Notified Body preparation.
- Regulatory gap assessments.
- Post-market surveillance activities.
- Temporary regulatory and quality resource support.
Established Global Manufacturers
Established medical device manufacturers often have experienced in-house regulatory and quality teams but may still require external specialist expertise, independent review or additional resources for complex or time-sensitive projects.
A medical device consultancy can work alongside internal teams to provide independent regulatory input, additional project capacity and specialist expertise in areas where additional support is required. This can be particularly valuable when entering new markets, managing significant product or manufacturing changes, addressing regulatory findings or reviewing established product portfolios.
Patient Guard works collaboratively with established manufacturers, adapting the level of support to complement existing regulatory and quality functions rather than replacing them.
Typical consultancy support includes:
- UK, EU and international market access.
- Regulatory remediation and gap assessments.
- Product portfolio and compliance reviews.
- Manufacturing and supplier transfers.
- Technical Documentation reviews and updates.
- Clinical and biological evaluation support.
- Regulatory change impact assessments.
- Independent regulatory due diligence.
- Notified Body and audit preparation.
- Post-market surveillance and ongoing compliance.
- Additional regulatory and quality project resources.
For established manufacturers, external consultancy can provide specialist expertise and additional capacity while allowing internal teams to retain ownership of their wider regulatory strategy and quality systems.
Software and Digital Health Companies
Software as a Medical Device (SaMD), medical device software, artificial intelligence and connected healthcare technologies can present regulatory challenges that differ significantly from those associated with traditional medical devices.
Software developers must first determine whether their product qualifies as a medical device and, where it does, establish the appropriate classification and regulatory pathway. Regulatory requirements must then be integrated into the software development lifecycle, rather than addressed only when the product is ready for market.
Medical device consultants can support software and digital health companies with requirements relating to:
- Medical device software qualification and classification.
- IEC 62304 software lifecycle processes.
- IEC 82304-1 for health software.
- IEC 62366-1 usability engineering.
- ISO 14971 risk management.
- Cybersecurity and information security.
- Clinical evaluation and clinical evidence.
- Artificial intelligence and machine learning considerations.
- EU MDR Rule 11 classification.
- UK medical device software requirements.
- Technical Documentation.
- Post-market surveillance and software change management.
- NHS DTAC and clinical safety requirements, where relevant to NHS adoption.
Whether developing standalone software, mobile applications, connected technologies or AI-enabled medical devices, Patient Guard can help organisations integrate regulatory, quality and clinical requirements throughout the software development lifecycle and establish an appropriate route to market.
In Vitro Diagnostic Manufacturers
Manufacturers of in vitro diagnostic medical devices must address the specific requirements of the EU In Vitro Diagnostic Medical Devices Regulation (IVDR), including device classification, performance evaluation, Technical Documentation, quality management and post-market performance monitoring.
The IVDR introduced a risk-based classification system for IVDs and places significant emphasis on demonstrating scientific validity, analytical performance and clinical performance throughout the lifecycle of the device.
Patient Guard supports IVD manufacturers with regulatory and quality activities including:
- IVDR classification and regulatory strategy.
- Technical Documentation preparation and review.
- Performance Evaluation Plans (PEPs).
- Scientific Validity Reports (SVRs).
- Analytical Performance Reports (APRs).
- Clinical Performance Reports (CPRs).
- Performance Evaluation Reports (PERs).
- Performance study planning and documentation.
- ISO 13485 Quality Management Systems.
- Risk management.
- Notified Body preparation and support.
- Post-Market Surveillance (PMS).
- Post-Market Performance Follow-up (PMPF).
- Performance Evaluation Report updates and lifecycle maintenance.
Whether developing a new IVD, transitioning an existing product or maintaining an established portfolio, specialist IVD consultancy can help manufacturers ensure that performance evidence, Technical Documentation and quality systems develop together throughout the device lifecycle.
Contract Manufacturers and OEMs
Contract manufacturers and Original Equipment Manufacturers (OEMs) can play an important role in the medical device supply chain, providing manufacturing, assembly, testing or other specialist activities on behalf of medical device manufacturers.
Where another organisation is the legal manufacturer, regulatory responsibilities must be clearly defined and the outsourced manufacturing activities appropriately controlled within the manufacturer’s Quality Management System. The legal manufacturer remains responsible for ensuring that the finished device complies with the applicable regulatory requirements.
Medical device consultants can support contract manufacturers, OEMs and their customers in establishing appropriate supplier controls, quality agreements, manufacturing documentation and change control processes, while ensuring that relevant manufacturing information is properly reflected within the device’s Technical Documentation and risk management activities.
Typical consultancy support includes:
- Supplier qualification and monitoring.
- ISO 13485 implementation and compliance.
- Supplier and contract manufacturer audits.
- Quality and technical agreements.
- Manufacturing process controls.
- Process validation requirements.
- Design and manufacturing transfers.
- Change control and regulatory impact assessment.
- Risk management.
- Traceability and manufacturing records.
- Technical Documentation support.
- Audit and Notified Body readiness.
Where regulatory roles are less straightforward, consultancy can also help organisations establish which entity is acting as the legal manufacturer and how responsibilities should be allocated between the organisations involved.
System and Procedure Pack Producers
System and procedure pack producers have specific regulatory responsibilities when combining CE-marked medical devices and, where applicable, other products for placement on the EU market under Article 22 of the EU MDR.
The regulatory pathway differs from conventional medical device manufacture. Organisations need to consider matters such as the compatibility of the combined products, verification of manufacturers’ instructions, appropriate packaging, information supplied with the system or procedure pack, registration requirements and the preparation of the required Article 22 statement.
Where devices are incorporated that do not bear the required CE marking, where combinations are incompatible with their original intended purposes, or where sterilisation activities fall outside the applicable Article 22 provisions, additional conformity assessment requirements may apply.
Patient Guard can support system and procedure pack producers with:
- Article 22 regulatory assessments.
- Review of system and procedure pack composition.
- Verification of device CE marking and regulatory status.
- Compatibility and intended-purpose assessments.
- Article 22 statements.
- Labelling and packaging reviews.
- Quality system and procedural requirements.
- Sterilisation considerations for applicable procedure packs.
- Registration and economic operator requirements.
- Supplier and manufacturer documentation reviews.
- Post-market and vigilance procedures.
- Regulatory gap assessments.
Importers and Distributors
Importers and distributors form an important part of the medical device supply chain and may have specific regulatory obligations depending on the market in which the device is being supplied and their role within that supply chain.
Under the EU MDR and IVDR, importers and distributors have defined responsibilities relating to verification, traceability, storage and transport, complaints and cooperation with manufacturers, Authorised Representatives and competent authorities. Requirements in Great Britain differ and should be assessed separately under the applicable UK medical device framework.
Patient Guard can help importers and distributors understand the obligations that apply to their activities and establish appropriate procedures within their quality systems to support ongoing compliance.
Typical consultancy support includes:
- Economic operator responsibility assessments.
- EU importer and distributor requirements.
- UK importer and distributor requirements.
- Verification of regulatory documentation and conformity status.
- Labelling and accompanying documentation reviews.
- Traceability and record-keeping procedures.
- Storage and transport controls.
- Complaint and vigilance procedures.
- Nonconforming device processes.
- Cooperation with manufacturers and regulatory authorities.
- Supplier and manufacturer due diligence.
- Regulatory and quality audits.
Specialist consultancy can also help organisations determine whether their activities could result in additional manufacturer-level regulatory responsibilities, particularly where devices are relabelled, repackaged, modified or supplied under another organisation’s name or trade mark.
Investors and Acquiring Organisations
Investors, private equity firms and organisations considering the acquisition of a medical device business may require independent regulatory due diligence before making an investment or completing a transaction.
A company’s regulatory position can have a significant effect on its commercial value. Deficiencies in Technical Documentation, clinical evidence, quality systems or post-market activities may create future remediation costs, delay market access or affect the continued commercialisation of existing products.
Patient Guard can provide an independent assessment of the organisation’s regulatory and quality position, helping investors and acquiring organisations understand potential compliance risks, documentation gaps and remediation requirements before completing a transaction.
Regulatory due diligence can include:
- Technical Documentation reviews.
- Quality Management System assessments.
- CE marking and UKCA compliance reviews.
- Device classification and regulatory pathway assessments.
- Clinical evidence and Clinical Evaluation reviews.
- Biological Evaluation reviews.
- Notified Body certification and audit finding reviews.
- Post-Market Surveillance and vigilance assessments.
- Regulatory gap analyses.
- Product portfolio evaluations.
- Pipeline and future market access assessments.
- Identification of potential regulatory remediation requirements.
The findings can provide investors and acquiring organisations with a clearer understanding of the regulatory maturity of the business, potential compliance liabilities and the work that may be required following investment or acquisition.
A Consultancy Partner That Grows With You
Every organisation faces different regulatory and quality challenges depending on its products, target markets, internal resources and stage of development.
Some manufacturers require specialist support for a single regulatory project, while others need additional capacity for their existing teams or choose to develop a longer-term consultancy relationship covering multiple products, markets and ongoing regulatory activities.
At Patient Guard, we tailor our medical device consultancy services to the specific needs of each organisation. Support can evolve as your business develops—from early regulatory strategy, classification and quality system implementation through to Technical Documentation, clinical and biological evaluation, market access, post-market surveillance and ongoing regulatory compliance.
Our aim is to provide practical, proportionate and commercially focused regulatory and quality support that complements your internal capabilities and evolves alongside your products and organisation.

Our Medical Device Consultancy Services
No two medical device manufacturers face exactly the same regulatory and quality challenges. Some organisations require support establishing their first Quality Management System, while others need specialist assistance with Technical Documentation, clinical or biological evaluation, responding to Notified Body findings or expanding into new international markets.
Patient Guard provides flexible medical device consultancy services across the medical device and IVD lifecycle. Our consultants can support individual regulatory or quality projects, provide specialist expertise in a particular area, or work as an extension of an existing regulatory affairs and quality team.
Our support can cover the journey from initial regulatory strategy and device classification through product development, quality management and Technical Documentation to market access, certification, post-market surveillance and ongoing regulatory compliance.
Whether you require support for a single device or an ongoing regulatory partnership across multiple products and markets, our approach is tailored to the device, regulatory pathway, target markets and internal capabilities of your organisation.
Regulatory Strategy
Every successful medical device regulatory project begins with a clear and practical strategy. Decisions made early in development can influence device classification, evidence requirements, quality management, Technical Documentation and the eventual route to market.
Our consultants help manufacturers establish an appropriate regulatory pathway based on the device, intended purpose, classification, target markets and stage of development. This allows regulatory requirements to be considered alongside product development and commercial planning rather than addressed retrospectively.
A well-defined regulatory strategy can help identify potential compliance issues early, avoid unnecessary testing or documentation, reduce the risk of costly development changes and provide a structured route towards market access.
Typical consultancy support includes:
- Medical device or IVD product determination.
- Intended purpose and claims review.
- Device classification.
- UK and EU regulatory pathway assessment.
- Regulatory roadmaps.
- Conformity assessment strategy.
- Identification of Notified Body requirements.
- Evidence and testing requirements.
- Regulatory gap assessments.
- Market access planning.
- Development and regulatory milestone planning.
Need Help Understanding Medical Device Classification?
Device classification is one of the first steps in developing an appropriate regulatory strategy. Learn how classification affects regulatory requirements and routes to market in our Guide to Medical Device Classification.
Planning Your Route to the UK Market?
Understand the regulatory framework for placing medical devices on the Great Britain market, including classification, UKCA marking, MHRA registration and UK Responsible Person requirements in our Complete Guide to UK Medical Device Regulations.
EU MDR and IVDR Compliance
The EU Medical Device Regulation (EU) 2017/745 (MDR) and In Vitro Diagnostic Medical Devices Regulation (EU) 2017/746 (IVDR) establish the regulatory frameworks for placing medical devices and IVDs on the European Union market.
Manufacturers must determine the appropriate device classification and conformity assessment route, demonstrate compliance with the applicable General Safety and Performance Requirements (GSPRs) and maintain sufficient technical, clinical or performance evidence to support the safety and performance of their devices.
Patient Guard supports medical device and IVD manufacturers throughout the regulatory lifecycle, from initial classification and regulatory strategy through Technical Documentation and quality management to conformity assessment, CE marking and ongoing post-market compliance.
Our consultancy services include:
- EU MDR compliance.
- EU IVDR compliance.
- Medical device and IVD classification.
- Regulatory strategy and conformity assessment planning.
- General Safety and Performance Requirements (GSPRs).
- Technical Documentation preparation and review.
- Clinical Evaluation for medical devices.
- Performance Evaluation for IVDs.
- ISO 13485 Quality Management Systems.
- Risk management.
- Notified Body preparation and support.
- EU Authorised Representative requirements.
- CE marking and Declaration of Conformity support.
- Post-Market Surveillance (PMS), PMCF and PMPF.
- Regulatory lifecycle and change management.
Need a Complete Guide to the EU MDR?
Understand the European Medical Device Regulation, including device classification, conformity assessment, Technical Documentation, clinical evaluation, CE marking and post-market requirements in our Complete Guide to the EU Medical Device Regulation (MDR) .
Need a Complete Guide to the EU IVDR?
Learn how the In Vitro Diagnostic Medical Devices Regulation affects IVD manufacturers, including classification, performance evaluation, Technical Documentation, conformity assessment and post-market requirements in our Complete Guide to the EU In Vitro Diagnostic Regulation (IVDR) .
UK Medical Device Regulations and UKCA
Manufacturers placing medical devices on the Great Britain market (England, Scotland and Wales) must comply with the applicable UK medical device regulatory framework. Medical devices in Great Britain are regulated by the Medicines and Healthcare products Regulatory Agency (MHRA) under the Medical Devices Regulations 2002 (SI 2002 No. 618, as amended). GOV.UK
The UKCA marking provides a UK conformity assessment route for medical devices. Depending on the device and regulatory route, qualifying CE-marked medical devices can also continue to be placed on the Great Britain market under the current transitional arrangements. GOV.UK
Patient Guard can help manufacturers understand the regulatory pathway applicable to their devices, prepare the necessary documentation and meet the requirements for placing and maintaining medical devices on the Great Britain market.
Our consultancy services include:
- UK medical device regulatory strategy.
- Device classification.
- UKCA conformity assessment requirements.
- Technical Documentation preparation and review.
- Quality Management System support.
- UK Approved Body preparation and support, where applicable.
- MHRA medical device registration.
- UK Responsible Person services for overseas manufacturers.
- Labelling and regulatory documentation reviews.
- Post-market surveillance and vigilance.
- Regulatory change management.
- Ongoing UK compliance support.
All medical devices placed on the Great Britain market must be registered with the MHRA. Manufacturers based outside the UK must appoint a UK Responsible Person to carry out specified responsibilities on their behalf.
Need a Complete Guide to UK Medical Device Regulations?
Understand the regulatory requirements for placing medical devices on the Great Britain market, including UKCA marking, MHRA registration, UK Responsible Persons, conformity assessment and the continued recognition of qualifying CE-marked devices in our Complete Guide to UK Medical Device Regulations .
Quality Management Systems
An effective Quality Management System (QMS) provides the framework through which medical device manufacturers control their regulatory and quality activities throughout the product lifecycle.
ISO 13485 is the internationally recognised quality management system standard for the medical device industry and provides a structured framework for areas including design and development, document control, supplier management, production, corrective and preventive action, internal auditing and continual improvement.
Patient Guard supports organisations with the implementation, maintenance and improvement of medical device Quality Management Systems, whether establishing a new QMS, preparing for ISO 13485 certification, addressing audit findings or improving an established quality system.
Our consultants can also work alongside existing quality teams to review individual processes, conduct independent audits and provide additional specialist support where required.
Typical consultancy services include:
- ISO 13485 QMS implementation.
- QMS gap assessments.
- Quality manuals, procedures and records.
- Document and record control.
- Design and development controls.
- Supplier qualification and management.
- Corrective and Preventive Action (CAPA).
- Nonconformity management.
- Internal audits.
- Supplier audits.
- Management Reviews.
- Risk-based quality processes.
- Certification and Notified Body audit preparation.
- QMS remediation and continual improvement.
- FDA Quality Management System Regulation (QMSR) support, where applicable.
Need a Complete Guide to ISO 13485?
Understand the quality management system requirements for medical device organisations, including implementation, documentation, risk-based processes, internal audits, management review and certification in our Complete Guide to ISO 13485 .
Technical Documentation
Technical Documentation provides the evidence that a medical device has been designed and developed to meet the applicable regulatory requirements and demonstrates how the manufacturer has addressed the device’s safety, performance and regulatory obligations.
Under the EU MDR, manufacturers must establish and maintain Technical Documentation in accordance with Annex II and Annex III, covering the device description, design and manufacturing information, General Safety and Performance Requirements (GSPRs), risk management, verification and validation, clinical evidence and post-market activities.
Patient Guard can support manufacturers with the preparation, review, remediation and ongoing maintenance of Technical Documentation, whether developing documentation for a new device, addressing gaps in an existing technical file or responding to findings raised during a Notified Body assessment.
Typical consultancy support includes:
- Technical Documentation preparation and review.
- EU MDR Annex II and Annex III documentation.
- Device description and specification.
- Intended purpose and classification documentation.
- General Safety and Performance Requirements (GSPR) checklists.
- Design and manufacturing information.
- Risk Management documentation.
- Verification and validation evidence.
- Clinical Evaluation documentation.
- Biological Evaluation documentation.
- Usability and software documentation, where applicable.
- Labelling and Instructions for Use (IFU) reviews.
- Post-Market Surveillance and PMCF documentation.
- Declaration of Conformity support.
- Technical Documentation gap assessments.
- Notified Body deficiency and remediation support.
- Lifecycle maintenance and change-related updates.
Building an EU MDR Technical File?
Learn how to structure robust Technical Documentation, avoid common deficiencies and prepare your medical device technical file for Notified Body review in our Guide to Building a Strong MDR Technical File .
Clinical Evaluation
Clinical evaluation is a fundamental part of demonstrating the safety, performance and clinical benefit of a medical device throughout its lifecycle.
Under the EU MDR, manufacturers must plan, conduct and document clinical evaluation in accordance with Article 61 and Annex XIV, using appropriate clinical data to demonstrate conformity with the relevant General Safety and Performance Requirements.
Patient Guard supports manufacturers throughout the clinical evaluation process, from establishing the clinical evaluation strategy and identifying relevant clinical evidence through to preparation and maintenance of the Clinical Evaluation Plan (CEP) and Clinical Evaluation Report (CER).
Our consultants can also help manufacturers identify gaps in existing clinical evidence and determine whether additional post-market or clinical activities may be required.
Typical consultancy support includes:
- Clinical Evaluation Plans (CEPs).
- Clinical Evaluation Reports (CERs).
- Clinical literature search strategies and protocols.
- Systematic literature searches.
- Clinical literature appraisal and analysis.
- State-of-the-art reviews.
- Evaluation of clinical data.
- Equivalence assessments, where applicable.
- Clinical evidence gap assessments.
- Post-Market Clinical Follow-up (PMCF) Plans.
- PMCF Evaluation Reports.
- Clinical evaluation lifecycle updates.
- Notified Body clinical evaluation remediation.
- Support with applicable MDCG guidance and MEDDEV 2.7/1 Rev. 4.
Need a Complete Guide to Medical Device Clinical Evaluation?
Understand Clinical Evaluation Plans, Clinical Evaluation Reports, clinical evidence, literature review, equivalence and PMCF under the EU MDR in our Complete Guide to Medical Device Clinical Evaluation .
Biological Evaluation
Demonstrating the biological safety of a medical device is an important part of the overall risk management process for devices that come into direct or indirect contact with the human body.
Biological evaluation should follow a risk-based approach based on the ISO 10993 series, considering factors such as the materials used, manufacturing processes, nature and duration of body contact, available biological and toxicological information and existing clinical experience.
Patient Guard supports manufacturers in developing and documenting an appropriate biological evaluation strategy. This includes assessing existing evidence, identifying potential biological risks and determining whether additional testing or other evidence is required to demonstrate biological safety.
Importantly, biological evaluation does not automatically mean that every biological endpoint requires new laboratory testing. Existing data, material information, chemical characterisation, toxicological evidence and scientific literature may contribute to the overall evaluation where appropriately justified.
Typical consultancy support includes:
- Biological Evaluation Plans (BEPs).
- Biological Evaluation Reports (BERs).
- ISO 10993-1 biological evaluation strategies.
- Biological risk and endpoint assessments.
- Material and device characterisation reviews.
- Existing biological and toxicological data reviews.
- Biocompatibility gap assessments.
- Identification and justification of testing requirements.
- Chemical characterisation strategy.
- Toxicological assessment support.
- Integration with ISO 14971 risk management.
- Evaluation of material, supplier and manufacturing changes.
- Review of laboratory test reports and existing evidence.
- Notified Body biological evaluation remediation.
Need a Complete Guide to Biological Evaluation?
Understand the risk-based approach to biological safety, including ISO 10993, Biological Evaluation Plans, Biological Evaluation Reports, biocompatibility evidence and testing strategies in our Complete Guide to Biological Evaluation of Medical Devices .
Risk Management
Risk management is a fundamental part of medical device development and should be maintained throughout the entire product lifecycle, from initial design and development through production, market use and post-market surveillance.
ISO 14971 provides the internationally recognised framework for applying risk management to medical devices. Manufacturers must establish a systematic process for identifying hazards, estimating and evaluating associated risks, implementing appropriate risk control measures and monitoring the effectiveness of those controls throughout the lifecycle of the device.
Patient Guard supports manufacturers with the development, review and maintenance of medical device risk management documentation, ensuring that risk management activities are appropriately integrated with Technical Documentation, clinical evaluation, biological evaluation, usability engineering, verification and validation and post-market surveillance.
Risk management should also evolve as new information becomes available. Complaints, vigilance data, post-market surveillance, design changes and emerging safety information should be assessed to determine whether existing risk assessments and control measures remain appropriate.
Typical consultancy support includes:
- ISO 14971 risk management strategies.
- Risk Management Plans.
- Hazard identification.
- Risk analysis and evaluation.
- Risk control measures.
- Evaluation of residual risk.
- Benefit-risk analysis, where required.
- Evaluation of overall residual risk.
- Risk Management Reports.
- Risk Management File reviews and gap assessments.
- Integration with clinical and biological evaluation.
- Integration with usability engineering.
- Production and post-production information.
- Post-market risk management updates.
- Change-related risk assessments.
- Notified Body risk management remediation.
Need a Complete Guide to ISO 14971 Risk Management?
Understand how to identify hazards, evaluate and control risks, assess residual risk and maintain risk management throughout the medical device lifecycle in our Complete Guide to ISO 14971 Medical Device Risk Management .
Medical Device Software
Medical device software can present distinct regulatory and quality challenges, particularly where software performs a medical purpose, forms part of a medical device or incorporates artificial intelligence or machine learning technologies.
Manufacturers must first determine whether the software qualifies as a medical device and, where applicable, establish the appropriate regulatory classification and conformity assessment route. Software regulatory requirements should then be integrated throughout the software development lifecycle, rather than addressed only when the product is ready for market.
Patient Guard supports manufacturers developing Software as a Medical Device (SaMD), embedded medical device software and digital health technologies, helping integrate regulatory, quality, risk management and clinical requirements into software development and maintenance processes.
Typical consultancy support includes:
- Medical device software qualification and classification.
- Software as a Medical Device (SaMD).
- IEC 62304 software lifecycle processes.
- Software safety classification.
- Software Development Plans.
- Software requirements and architecture documentation.
- Software verification and validation.
- ISO 14971 software risk management.
- IEC 62366-1 usability engineering.
- Cybersecurity and information security considerations.
- Software configuration and change management.
- Clinical evaluation for medical device software.
- Technical Documentation.
- EU MDR Rule 11 classification considerations.
- UK medical device software requirements.
- Artificial intelligence and machine-learning considerations.
- Post-market surveillance and software lifecycle maintenance.
For medical device software, compliance should develop alongside the product. Establishing appropriate lifecycle, risk management, usability and verification processes early can help manufacturers build the evidence required to demonstrate the safety and performance of the software throughout its lifecycle.
Need a Complete Guide to IEC 62304?
Understand the medical device software lifecycle, including software safety classification, development planning, architecture, verification, maintenance and risk management in our Complete Guide to IEC 62304 Medical Device Software Development .
Validation Services
Validation provides documented evidence that processes, systems and methods consistently achieve their intended results and is an important component of medical device quality management and regulatory compliance.
Depending on the device and manufacturing process, manufacturers may need to validate activities where the resulting output cannot be fully verified through subsequent inspection or testing. Validation requirements can also apply to areas such as sterilisation, cleaning, packaging, transportation, software and computerised systems.
Patient Guard supports manufacturers in developing appropriate validation strategies, plans, protocols and reports, as well as reviewing existing validation evidence to identify gaps against applicable regulatory requirements, standards and defined acceptance criteria.
Our role can include defining and documenting validation requirements and reviewing evidence generated by manufacturers, suppliers and specialist laboratories.
Typical consultancy support includes:
- Validation strategy and planning.
- Process validation.
- Cleaning validation.
- Sterilisation validation.
- Packaging validation.
- Transportation and distribution validation.
- Computer system validation.
- Software validation, where applicable.
- Test method validation.
- Validation Plans and protocols.
- Definition and review of acceptance criteria.
- Validation Reports.
- Review of supplier and laboratory validation evidence.
- Validation gap assessments.
- Integration of validation evidence into Technical Documentation.
- Validation remediation and Notified Body support.
Need a Complete Guide to Medical Device Validation?
Understand medical device validation requirements, including process, cleaning, sterilisation, packaging and other validation activities in our Complete Guide to Validation of Medical Devices .
Notified Body Support
Preparing for a Notified Body assessment requires careful planning, robust Technical Documentation and an effective Quality Management System. Identifying potential compliance gaps before formal assessment can help manufacturers address weaknesses early and reduce the risk of avoidable findings or delays.
Patient Guard supports manufacturers before, during and after Notified Body conformity assessment activities, providing independent regulatory and quality expertise across Technical Documentation, clinical and biological evidence, risk management, quality systems and post-market requirements.
Our consultants can conduct gap assessments and independent reviews to identify areas requiring attention before submission or audit. Where a Notified Body has already raised findings or questions, we can also support manufacturers in determining root causes, developing corrective actions and preparing appropriate remediation.
Typical consultancy support includes:
- Notified Body readiness assessments.
- Regulatory and QMS gap assessments.
- Technical Documentation reviews.
- ISO 13485 internal audits.
- Mock Notified Body audits.
- Clinical Evaluation reviews.
- Biological Evaluation reviews.
- Risk Management File reviews.
- Post-Market Surveillance and PMCF reviews.
- Audit preparation and staff support.
- Review of Notified Body findings and deficiencies.
- Root cause analysis.
- Corrective and Preventive Action (CAPA).
- Technical Documentation remediation.
- Support with responses to Notified Body questions.
- Surveillance and recertification preparation.
- Ongoing regulatory remediation and compliance support.
Need a Complete Guide to Notified Bodies?
Understand the role of Notified Bodies under the EU MDR and IVDR, including conformity assessment, Technical Documentation review, quality system assessment and ongoing surveillance in our Complete Guide to Medical Device Notified Bodies .
Post-Market Surveillance
Regulatory compliance continues throughout the entire medical device lifecycle. Once a device is placed on the market, manufacturers must systematically collect, review and evaluate information relating to its safety, performance and use.
An effective Post-Market Surveillance (PMS) system enables manufacturers to identify emerging risks, detect trends, evaluate real-world device performance and determine whether corrective or preventive action may be required.
Patient Guard supports manufacturers in developing, implementing and maintaining appropriate post-market surveillance processes and documentation. Information generated through PMS should also feed back into other regulatory activities, including risk management, clinical evaluation, Technical Documentation and the Quality Management System.
Typical consultancy support includes:
- Post-Market Surveillance Plans.
- PMS Reports.
- Periodic Safety Update Reports (PSURs).
- Post-Market Clinical Follow-up (PMCF) Plans.
- PMCF Evaluation Reports.
- Complaint and feedback data analysis.
- Vigilance and incident assessment.
- Trend reporting.
- Literature surveillance.
- Signal and trend analysis.
- Review of post-market safety and performance data.
- Integration of PMS findings with Clinical Evaluation.
- Risk Management File updates.
- Technical Documentation updates.
- Corrective and Preventive Action (CAPA) support.
- PMS gap assessments and remediation.
- Ongoing lifecycle maintenance of post-market documentation.
For IVD manufacturers, Patient Guard can also support Post-Market Performance Follow-up (PMPF) and the integration of post-market performance information into the device’s Performance Evaluation.
Need a Complete Guide to Post-Market Surveillance?
Understand medical device post-market requirements, including PMS Plans, PMS Reports, PSURs, PMCF, vigilance, trend reporting and the use of post-market data throughout the device lifecycle in our Complete Guide to Medical Device Post-Market Surveillance .
Regulatory Training
Building internal regulatory and quality capability is an important part of maintaining long-term medical device compliance. Appropriate training helps employees understand their responsibilities, apply regulatory and quality requirements consistently and maintain the competence required for their roles.
Patient Guard provides practical regulatory and quality training for medical device and IVD organisations, covering international regulations, standards and quality management requirements.
Training can be delivered to individual teams or across an organisation and tailored to the products, regulatory markets, Quality Management System and responsibilities of the employees involved.
Training topics can include:
- EU Medical Device Regulation (MDR).
- EU In Vitro Diagnostic Regulation (IVDR).
- UK medical device regulations.
- ISO 13485 Quality Management Systems.
- ISO 14971 medical device risk management.
- ISO 9001 Quality Management Systems.
- ISO/IEC 27001 Information Security Management Systems.
- Technical Documentation.
- Clinical Evaluation.
- Biological Evaluation.
- Post-Market Surveillance and PMCF/PMPF.
- IEC 62304 medical device software.
- IEC 62366-1 usability engineering.
- Internal auditing.
- Regulatory awareness.
- Quality Management System processes and responsibilities.
- Bespoke regulatory and quality training.
Training can be delivered remotely or on-site and adapted to the organisation’s existing level of regulatory knowledge, helping ensure that the content is relevant to the employees and activities involved.
Looking for Medical Device Regulatory and Quality Training?
Explore our practical training covering medical device regulations, quality management systems, ISO standards and regulatory compliance, with courses available remotely or tailored to your organisation's requirements. View Patient Guard Training Courses .

Why Choose Patient Guard?
Choosing the right medical device consultancy is about more than finding regulatory expertise. Manufacturers need a partner who understands their technology, communicates clearly and can translate complex regulatory requirements into practical and proportionate actions.
At Patient Guard, we specialise in medical device and IVD regulatory affairs and quality assurance, supporting manufacturers throughout the entire product lifecycle — from early regulatory strategy and product development through Technical Documentation, quality management and conformity assessment to post-market surveillance and ongoing compliance.
Our multidisciplinary team brings together expertise across medical device regulation, quality management, clinical evaluation, biological evaluation, risk management, software, validation and post-market activities. This allows different areas of regulatory compliance to be considered together rather than treated as isolated projects.
We work collaboratively with manufacturers, adapting our support to their existing capabilities and requirements. This may involve completing a single specialist project, providing additional expertise to an established regulatory or quality team, or developing a longer-term consultancy relationship across multiple products and markets.
Our approach is focused on providing clear, practical and commercially aware regulatory support, helping manufacturers identify compliance risks, make informed regulatory decisions and establish a structured route towards UK and EU market access and ongoing compliance.
Comprehensive Regulatory Expertise
Medical device regulation continues to evolve across the UK, European Union and international markets, requiring manufacturers to manage increasingly interconnected regulatory, quality, clinical and technical requirements.
Patient Guard brings together expertise across multiple medical device and IVD disciplines, allowing manufacturers to access specialist regulatory and quality knowledge through a single consultancy partner.
Our expertise includes:
- EU MDR (Regulation (EU) 2017/745).
- EU IVDR (Regulation (EU) 2017/746).
- UK medical device regulations.
- ISO 13485 Quality Management Systems.
- ISO 14971 Risk Management.
- ISO 10993 Biological Evaluation.
- Clinical and Performance Evaluation.
- Technical Documentation.
- IEC 62304 medical device software.
- IEC 62366-1 usability engineering.
- Medical device validation.
- Post-Market Surveillance, PMCF and PMPF.
- Notified Body and audit preparation.
- Regulatory strategy and market access.
These disciplines do not operate independently. Risk management informs clinical and biological evaluation, validation evidence supports Technical Documentation, and post-market information feeds back into clinical evaluation, risk management and ongoing regulatory compliance.
By considering these requirements together, Patient Guard can support manufacturers across the entire regulatory lifecycle, while helping ensure that different elements of the regulatory strategy, Technical Documentation and Quality Management System remain aligned as the device develops.
Practical, Commercial Advice
Regulatory compliance should support the development of safe, effective and commercially viable medical devices, rather than create unnecessary barriers to innovation.
Patient Guard understands that manufacturers operate within real-world commercial constraints, including development budgets, product launch timelines, available internal resources and investor expectations. Regulatory strategies therefore need to be technically robust while remaining practical and proportionate to the device and its associated risks.
Our consultants focus on identifying what is actually required, making appropriate use of existing evidence where possible and avoiding unnecessary regulatory activities that do not add meaningful value to the compliance strategy.
By considering regulatory requirements alongside product development and commercial objectives, we help manufacturers prioritise activities, allocate resources effectively and establish a proportionate route towards market access, without compromising the regulatory requirements applicable to the device.
Support Throughout the Entire Product Lifecycle
Regulatory requirements change as a medical device progresses from initial concept through development, market access and post-market use. Working with a consultancy that understands the product throughout this journey can provide valuable continuity in regulatory and quality decision-making.
Patient Guard can support manufacturers from early regulatory strategy and device classification through product development, quality management, Technical Documentation and conformity assessment to post-market surveillance and ongoing compliance.
This continuity allows regulatory decisions made during the early stages of development to remain connected with the evidence and documentation generated later in the lifecycle. It can also help maintain alignment between risk management, clinical and biological evaluation, validation, Technical Documentation, the Quality Management System and post-market activities.
As products, markets and regulatory requirements develop, our level of support can evolve alongside the manufacturer — from a single specialist project to additional capacity for an internal team or an ongoing regulatory and quality partnership.
By maintaining this continuity, manufacturers can identify potential compliance issues earlier, reduce unnecessary duplication and keep regulatory activities aligned throughout the product lifecycle.
Flexible Consultancy That Fits Your Business
Every medical device manufacturer has different regulatory and quality requirements, and the level of external support needed can change throughout the product lifecycle.
Some clients require Patient Guard’s expertise for a single defined project, such as a Clinical Evaluation Report, Biological Evaluation or Technical Documentation review. Others require additional capacity for an established regulatory or quality team, while some choose an ongoing consultancy relationship covering multiple products, regulatory disciplines and international markets.
Patient Guard can provide flexible support as:
- Project-based regulatory and quality consultancy.
- Ongoing retained regulatory support.
- An extension of an existing QA/RA team.
- Independent technical and regulatory reviewers.
- Specialist expertise for defined regulatory activities.
- Temporary regulatory or quality resource during periods of increased workload or staff absence.
- Longer-term support across multiple products and markets.
This approach allows manufacturers to access experienced regulatory and quality professionals when they are needed, without necessarily increasing permanent headcount or outsourcing their entire regulatory function.
As an organisation grows, launches new products or enters additional markets, the level and type of consultancy support can evolve alongside the business, providing additional expertise and capacity while allowing the manufacturer’s internal team to retain ownership of its regulatory and quality activities.
Quality, Communication and Transparency
Successful regulatory consultancy depends on more than technical expertise. Clear communication, defined responsibilities and effective project management are equally important to delivering regulatory and quality projects efficiently.
At Patient Guard, we aim to establish clear expectations from the outset, including the scope of work, deliverables, responsibilities, timescales and costs associated with each project. As work progresses, we communicate significant findings, potential regulatory issues and any information or actions required from the manufacturer.
Our clients should always understand:
- What work is being undertaken.
- Why the work is required.
- What information or input is needed from them.
- The current status of the project.
- Expected deliverables and timescales.
- Any significant regulatory or quality issues identified.
- The potential implications of those issues.
- Any recommended next steps.
Quality is also embedded within our own operations. Patient Guard operates an ISO 13485-certified Quality Management System, providing a structured framework for the control and delivery of our medical device regulatory and quality services.
By combining technical expertise with clear communication, transparent project management and controlled quality processes, we aim to give manufacturers the information they need to make informed regulatory decisions throughout each project.
A Long-Term Regulatory Partner
Medical device compliance does not end once a product reaches the market. Manufacturers must continue to maintain their Technical Documentation, clinical and biological evidence, Quality Management System, risk management and post-market activities throughout the commercial lifecycle of the device.
Many manufacturers continue working with Patient Guard after their initial project, giving them ongoing access to regulatory and quality professionals who already understand their devices, documentation, regulatory strategy and quality systems.
As products evolve, we can support changes to existing devices, new product development, entry into additional markets, Technical Documentation and Clinical Evaluation updates, biological safety reviews, internal and supplier audits, post-market activities and regulatory change assessments.
This continuity can be particularly valuable as a manufacturer’s product portfolio grows. Rather than approaching each regulatory activity as a separate project, Patient Guard can help ensure that new information, product changes and regulatory developments are considered across the wider regulatory and quality framework.
Our aim is to develop long-term relationships that give manufacturers access to consistent regulatory and quality expertise as their requirements evolve, from initial development and market access through to established products and ongoing post-market compliance.

Peter Reeve
Director
Trusted by Medical Device Manufacturers
Patient Guard supports organisations across the medical device and IVD sector, from innovative start-ups developing their first regulated product to established international manufacturers managing complex product portfolios and multiple regulatory markets.
Since Patient Guard was established in 2017, we have supported more than 500 customers with regulatory affairs and quality assurance activities across the medical device and IVD lifecycle.
Our regulatory and quality specialists work across disciplines including Technical Documentation, Quality Management Systems, Clinical and Performance Evaluation, Biological Evaluation, Risk Management, medical device software, validation and post-market surveillance, enabling manufacturers to access specialist expertise through a single consultancy partner.
Patient Guard also operates an ISO 13485-certified Quality Management System, applying structured quality processes to the delivery and control of our own medical device regulatory and quality services.
Clients can engage Patient Guard for a single specialist project, additional support for an existing regulatory or quality team, or a longer-term regulatory partnership as their products, markets and organisations develop.
Whether the objective is preparing for CE marking, entering the Great Britain market, strengthening a Quality Management System, preparing for Notified Body assessment or maintaining established products, our approach is centred on practical, proportionate and clearly communicated regulatory support.


Ready to Discuss Your Project?
Whether you require support with a single regulatory or quality project, additional expertise for your existing team or an ongoing consultancy partnership across your product portfolio, Patient Guard can help.
Our consultants will take the time to understand your device, current stage of development, target markets and regulatory challenges before recommending a practical and proportionate approach to the support you require.
From early regulatory strategy and product development through to market access and ongoing compliance, our medical device consultancy services can be tailored to the needs of your organisation.
Contact Patient Guard today to discuss your medical device project and the next steps towards regulatory compliance and market access.

Our Medical Device Consultancy Process
Every medical device project is different. The appropriate regulatory approach depends on factors including the device, intended purpose, classification, stage of development, target markets and manufacturer’s existing regulatory and quality capabilities.
At Patient Guard, we begin by understanding the product, the organisation’s objectives and the regulatory or quality challenges that need to be addressed. This allows us to establish a practical and proportionate scope of work based on the requirements applicable to the project.
Our consultancy process is designed to provide clear responsibilities, defined deliverables, realistic timescales and transparent communication throughout the project, while allowing our consultants to work collaboratively with the manufacturer’s internal teams and other relevant specialists.
Whether you require support with a single Technical Documentation review, a specialist Clinical or Biological Evaluation project, implementation of a Quality Management System or a broader regulatory strategy covering multiple products and markets, the same structured approach is applied.
The objective is to give manufacturers a clear understanding of what needs to be done, why it is required and how each activity contributes to the wider regulatory pathway, while maintaining flexibility as the project develops.
Step 1: Discovery and Initial Consultation
Every project begins with a discussion to understand the device, organisation, regulatory position and objectives before defining the scope of consultancy support.
During this initial stage, we seek to understand:
- The device, technology and intended purpose.
- Current stage of product development.
- Existing regulatory status and device classification, where established.
- Applicable or intended regulatory markets.
- Existing Technical Documentation and supporting evidence.
- Current Quality Management System arrangements.
- Regulatory or quality challenges already identified.
- Target markets.
- Internal regulatory and quality resources.
- Commercial objectives and priorities.
- Project milestones and anticipated timescales.
This initial assessment allows Patient Guard to understand the wider regulatory context and identify where specialist support is genuinely required, rather than applying a standard package of services to every project.
From this information, we can begin to establish an appropriate regulatory strategy, project scope and priorities before substantive consultancy work begins.
Step 2: Regulatory Gap Assessment
Where appropriate, Patient Guard can review the manufacturer’s existing regulatory documentation, evidence and quality systems to assess the current position against the requirements applicable to the device and target market.
The scope of the assessment will depend on the project but may include:
- Device classification and regulatory pathway.
- Technical Documentation.
- Quality Management System documentation.
- Risk Management.
- Clinical or Performance Evaluation.
- Biological Evaluation.
- Verification and validation evidence.
- Software and usability documentation, where applicable.
- Labelling and Instructions for Use.
- Post-Market Surveillance, PMCF or PMPF.
- Supplier and outsourced process controls.
- Existing certification and conformity assessment documentation.
The assessment can identify missing documentation, insufficient evidence, inconsistencies between regulatory documents and areas requiring further development or remediation. Existing evidence can also be reviewed to determine what may remain suitable and where updates or additional work may be required.
The outcome is a prioritised action plan that distinguishes the most significant regulatory issues from lower-priority improvements, helping the manufacturer focus resources on the activities that are most important to progressing the regulatory pathway.
Step 3: Developing Your Regulatory Strategy
Following the initial assessment, Patient Guard can develop a structured regulatory roadmap based on the device, current compliance position, target markets and manufacturer’s objectives.
The strategy defines the regulatory activities required to progress the device towards market access and helps establish the appropriate sequence and priorities for completing them.
Depending on the project, this may include:
- Device qualification and classification.
- Regulatory pathway and conformity assessment strategy.
- Identification of applicable regulations and standards.
- Quality Management System requirements.
- Technical Documentation requirements.
- Risk management strategy.
- Verification and validation requirements.
- Testing strategy.
- Clinical or Performance Evaluation planning.
- Biological Evaluation strategy, where applicable.
- Software and usability requirements, where applicable.
- Notified Body or UK Approved Body involvement, where required.
- EU Authorised Representative or UK Responsible Person requirements, where applicable.
- Registration and market access requirements.
- Post-Market Surveillance requirements.
- Project priorities, dependencies and anticipated milestones.
Where appropriate, existing documentation, testing and evidence can be considered as part of the strategy to determine what may remain suitable, what requires updating and where additional evidence or activities are needed.
The outcome is a practical regulatory roadmap that gives the manufacturer a clearer understanding of what needs to be completed, in what order and how the different regulatory activities fit together, providing a structured route towards market access and ongoing compliance.
Step 4: Documentation and Implementation
Once the regulatory strategy and scope of work have been agreed, Patient Guard can begin the preparation, review or remediation of the documentation and systems required to support regulatory compliance.
Depending on the project, our consultants may work directly alongside the manufacturer’s regulatory, quality, engineering and clinical teams, using available product information and existing evidence while identifying any additional information or activities required.
Typical activities may include:
- Technical Documentation preparation and review.
- Clinical Evaluation Plans and Reports.
- Biological Evaluation Plans and Reports.
- Risk Management documentation.
- Quality Management System implementation and documentation.
- Verification and validation documentation.
- Software and usability documentation, where applicable.
- Labelling and Instructions for Use reviews.
- Post-Market Surveillance and PMCF/PMPF documentation.
- Notified Body or UK Approved Body preparation, where applicable.
- Regulatory registration and market access documentation.
- Remediation of existing regulatory or quality documentation.
Throughout delivery, we maintain clear communication with the manufacturer’s team, providing progress updates, identifying outstanding information or actions and discussing significant regulatory issues as they arise.
Our collaborative approach allows responsibilities to be divided appropriately between Patient Guard and the manufacturer, ensuring that internal expertise and existing evidence are used effectively while specialist regulatory and quality support is provided where required.
Step 5: Conformity Assessment and Market Access
Once the required regulatory documentation, evidence and quality systems are in place, the manufacturer can progress through the applicable conformity assessment and market access activities for the target market.
Where third-party conformity assessment is required, Patient Guard can support manufacturers in preparing for and navigating interactions with EU Notified Bodies or UK Approved Bodies. The precise route will depend on factors including the device classification, regulatory framework and target market.
Depending on the project, support may include:
- Notified Body or UK Approved Body readiness assessments.
- Technical Documentation review before assessment.
- Quality Management System and audit preparation.
- Support during conformity assessment activities.
- Responses to technical or regulatory questions.
- Review and remediation of assessment findings.
- Root cause analysis and CAPA support.
- CE marking and UKCA conformity assessment support.
- Declaration of Conformity documentation.
- Regulatory registration requirements.
- EU Authorised Representative or UK Responsible Person requirements, where applicable.
- Regulatory authority interactions, where required.
For devices that do not require third-party conformity assessment, Patient Guard can support manufacturers in identifying and completing the applicable self-declaration, documentation and registration requirements before placing the device on the relevant market.
Our objective is to help manufacturers enter the assessment or market access process with appropriate documentation, evidence and quality systems in place, while providing regulatory support where questions, findings or additional requirements arise.
Step 6: Ongoing Regulatory Support
Regulatory compliance continues throughout the commercial lifecycle of a medical device. Once a product has been placed on the market, manufacturers must continue to maintain their regulatory documentation, monitor device performance and respond appropriately to new information, product changes and evolving regulatory requirements.
Many organisations continue working with Patient Guard after initial market access, providing continuity and access to regulatory and quality professionals who already understand their devices, documentation, quality systems and regulatory strategy.
Ongoing consultancy support may include:
- Technical Documentation maintenance and updates.
- Clinical Evaluation updates.
- Biological Evaluation reviews and updates.
- Risk Management File updates.
- Post-Market Surveillance.
- PMCF and PMPF activities, where applicable.
- Vigilance and complaint-related regulatory support.
- Internal audits.
- Supplier audits.
- Quality Management System support.
- Product and manufacturing change assessments.
- Regulatory change management.
- New product development and market expansion.
- Preparation for surveillance and recertification activities.
Support can be provided for individual activities as they arise or through a longer-term regulatory and quality consultancy relationship, depending on the manufacturer’s internal resources and requirements.
By maintaining continuity throughout the device lifecycle, Patient Guard can help manufacturers identify and respond to regulatory changes, emerging risks and product developments while maintaining the documentation, evidence and systems required to support continued market access.

A Partnership Built Around Your Business
Every organisation has different products, objectives, internal resources and regulatory challenges, so the level of consultancy support required will vary from one manufacturer to another.
Some clients engage Patient Guard for a single specialist project, while others work with us across multiple regulatory activities or use our consultants as an extension of their internal regulatory affairs and quality teams.
Our approach is designed to complement your existing capabilities. We can take responsibility for defined regulatory workstreams, provide specialist expertise where it is not available internally, or add additional capacity during periods of increased workload.
Whatever the size or complexity of the project, our focus remains the same: practical and proportionate regulatory advice, clearly defined responsibilities, transparent communication and specialist expertise tailored to your organisation and products.
This flexible approach allows Patient Guard to support your organisation as its requirements evolve, from an individual regulatory project through to an ongoing regulatory and quality partnership.

🏆 Patient Guard's BSI ISO 13485 Certification
Choosing a regulatory consultancy that operates under its own certified Quality Management System provides additional confidence that your compliance partner follows the same high standards expected of medical device manufacturers. Patient Guard maintains a BSI-certified ISO 13485:2016 Quality Management System, independently audited to demonstrate our commitment to quality, regulatory compliance and continual improvement. View our certification and learn what it means for our clients.
Frequently Asked Questions About Medical Device Consultancy
A medical device consultancy provides specialist regulatory, quality assurance and technical expertise to organisations developing, manufacturing or marketing medical devices and in vitro diagnostic (IVD) devices. Consultants help manufacturers understand regulatory requirements, prepare technical documentation, implement quality management systems, achieve market approvals and maintain compliance throughout the product lifecycle.
The best time to engage a medical device consultancy is as early as possible during product development. Early regulatory planning helps determine device classification, identify applicable regulations, develop a realistic route to market and avoid costly redesigns or compliance issues later in the project. Consultants can also provide valuable support during market expansion, regulatory remediation and post-market compliance.
Medical device consultancies typically provide support across a wide range of regulatory and quality activities, including regulatory strategy, device classification, Technical Documentation, Clinical Evaluation, Biological Evaluation, ISO 13485 Quality Management Systems, ISO 14971 Risk Management, Software as a Medical Device (SaMD), validation, CE marking, UKCA compliance, Post-Market Surveillance and Notified Body support.
Yes. Medical device consultancies frequently support manufacturers preparing for CE marking by developing Technical Documentation, Clinical Evaluation Reports, Risk Management Files, Quality Management Systems and other documentation required under the EU Medical Device Regulation (EU MDR) or In Vitro Diagnostic Regulation (IVDR). Consultants can also assist with Notified Body preparation and regulatory submissions where applicable.
Yes. Patient Guard supports manufacturers placing products on the Great Britain market by providing consultancy covering UK Medical Device Regulations, UKCA marking, UK Responsible Person services, MHRA registration requirements and ongoing regulatory compliance.
Absolutely. Many of our clients are start-ups developing their first medical device. We help founders establish regulatory strategies, determine device classifications, implement Quality Management Systems, prepare technical documentation and develop realistic routes to market that support both regulatory success and commercial growth.
Many medical device manufacturers require a Quality Management System aligned with ISO 13485 to demonstrate compliance with regulatory requirements and support conformity assessment. Even where certification is not mandatory, implementing ISO 13485 provides a structured framework for quality, risk management and continual improvement that benefits manufacturers throughout the product lifecycle.
Yes. We prepare and review Technical Documentation for medical devices and IVDs in accordance with applicable regulatory requirements. This includes device descriptions, General Safety and Performance Requirements (GSPR) checklists, Clinical Evaluation, Biological Evaluation, Risk Management documentation, labelling reviews and supporting evidence required for conformity assessment.
Yes. Patient Guard provides comprehensive support for Clinical Evaluation Plans (CEPs), Clinical Evaluation Reports (CERs), Biological Evaluation Plans (BEPs) and Biological Evaluation Reports (BERs). Our consultants help manufacturers develop evidence-based documentation aligned with current MDR, IVDR and ISO 10993 requirements.
Yes. We regularly assist manufacturers preparing for initial certification, surveillance audits and Technical Documentation reviews conducted by Notified Bodies. Our support includes gap assessments, internal audits, documentation reviews, CAPA implementation and responses to regulatory findings.
Yes. Regulatory compliance continues after products reach the market. Patient Guard supports manufacturers with Post-Market Surveillance, PMCF, vigilance, Technical Documentation updates, Clinical Evaluation updates, internal audits, supplier management and regulatory change assessments throughout the commercial life of their products.
Yes. Our consultants support manufacturers developing medical device software, digital health technologies and AI-enabled medical devices. We provide guidance covering IEC 62304, IEC 62366-1, software classification, cybersecurity, clinical evidence and software lifecycle documentation.
Yes. Many organisations use Patient Guard as an extension of their internal regulatory and quality teams. We provide flexible consultancy ranging from individual specialist projects through to fully outsourced Regulatory Affairs and Quality Assurance support, allowing manufacturers to access experienced professionals without expanding their permanent workforce.
We support manufacturers across a broad range of healthcare technologies, including medical devices, in vitro diagnostics, Software as a Medical Device (SaMD), digital health technologies, artificial intelligence, dental devices, ophthalmic devices, wound care products, implantable devices, diagnostics and many other specialist medical technologies.
Patient Guard combines extensive regulatory expertise with practical commercial experience to help manufacturers achieve compliance efficiently and confidently. Our consultants provide tailored support across the entire product lifecycle, from regulatory strategy and product development through to Technical Documentation, Quality Management Systems, market approval and ongoing regulatory compliance. Whether you require support for a single project or a long-term regulatory partnership, we work collaboratively with your team to deliver practical, proportionate and effective regulatory solutions.
The cost of medical device consultancy depends on the complexity of your project, the type of device, the target markets and the level of support required. A simple regulatory review or gap assessment may require only a few days of consultancy, whereas developing complete Technical Documentation, implementing an ISO 13485 Quality Management System or preparing for a Notified Body assessment will require a more comprehensive project. At Patient Guard, we provide transparent quotations tailored to your specific requirements, ensuring you understand the scope of work and associated costs before the project begins.
Regulatory timelines vary considerably depending on the classification of your device, the regulatory pathway, the quality of existing documentation and whether third-party assessment is required. Some projects, such as regulatory strategy reviews or gap assessments, can be completed within days or weeks, while preparing complete Technical Documentation and obtaining CE marking may take several months. Engaging an experienced medical device consultancy early in the development process can help identify potential delays, streamline documentation and reduce the overall time required to achieve market access.
Yes. Patient Guard works with medical device and IVD manufacturers around the world, supporting organisations seeking access to the UK, European Union and other international markets. Our consultants regularly assist companies with EU MDR, EU IVDR and UK Medical Device Regulations, as well as providing services including Technical Documentation, Clinical Evaluation, Biological Evaluation, Quality Management Systems, Regulatory Strategy and market access planning. Whether you are an established international manufacturer or an overseas company entering new markets for the first time, we can provide practical regulatory support tailored to your products and business objectives.
References
This guide is based on the principal UK and European medical device regulations, internationally recognised medical device standards and current regulatory guidance relevant to medical device consultancy, quality management, risk management, technical documentation, clinical and biological evaluation, software, conformity assessment and post-market compliance.
| Organisation | Reference | Why it's relevant |
|---|---|---|
| European Union | Regulation (EU) 2017/745 – Medical Device Regulation (MDR) | Establishes the European regulatory framework for medical devices, including manufacturer obligations, classification, conformity assessment, Technical Documentation, clinical evaluation, economic operators, post-market surveillance and vigilance. |
| European Union | Regulation (EU) 2017/746 – In Vitro Diagnostic Medical Devices Regulation (IVDR) | Establishes the European regulatory framework for in vitro diagnostic medical devices, including risk-based classification, performance evaluation, Technical Documentation, conformity assessment and post-market performance requirements. |
| European Commission | Medical Device Coordination Group (MDCG) Guidance | Provides European guidance supporting implementation of the MDR and IVDR across areas including classification, clinical and performance evaluation, medical device software, post-market surveillance, vigilance and economic operator responsibilities. |
| UK Government / MHRA | MHRA – Regulating Medical Devices in the UK | Provides guidance on placing medical devices on the UK market, including UKCA marking, recognition of qualifying CE-marked devices, MHRA registration, UK Responsible Persons, UK Approved Bodies and post-market requirements. |
| UK Government | Medical Devices Regulations 2002 (SI 2002 No. 618), as amended | Provides the legislative framework governing medical devices in Great Britain and establishes applicable requirements for placing devices on the market and meeting ongoing regulatory obligations. |
| ISO | ISO 13485:2016 – Medical Devices Quality Management Systems | Specifies quality management system requirements for medical device organisations, including document control, design and development, supplier management, production controls, CAPA, internal auditing and continual improvement. |
| ISO | ISO 14971:2019 – Application of Risk Management to Medical Devices | Provides the internationally recognised framework for identifying hazards, estimating and evaluating risks, implementing risk controls and monitoring risk throughout the medical device lifecycle. |
| ISO | ISO 10993-1:2025 – Biological Evaluation of Medical Devices | Establishes requirements and general principles for evaluating the biological safety of medical devices within a risk management process, including consideration of biological hazards, exposure, material characterisation and biological safety evidence. |
| IEC | IEC 62304:2006 – Medical Device Software Lifecycle Processes | Defines lifecycle processes for medical device software, including software development, maintenance, configuration management, problem resolution and software risk-related activities. |
| IEC | IEC 62366-1 – Usability Engineering for Medical Devices | Establishes a usability engineering process for medical devices and supports the identification, evaluation and control of risks associated with device use and user interaction. |
| European Commission | MDCG Guidance on Medical Device Classification | Provides guidance on applying the classification rules established by the MDR and supports manufacturers in determining the appropriate device classification and conformity assessment pathway. |
| European Commission | MDCG Guidance on Medical Device Software | Provides guidance relating to the qualification and classification of medical device software and supports interpretation of regulatory requirements applicable to software and digital health technologies. |
| European Commission | MDCG Guidance on Post-Market Surveillance and Vigilance | Supports manufacturers in interpreting and implementing post-market surveillance, vigilance, trend reporting and related lifecycle requirements under the MDR and IVDR. |
Medical device regulations, standards and regulatory guidance are periodically amended or updated. Manufacturers should verify that they are working to the current regulatory requirements, consolidated legislation, standards and applicable guidance for their device, target market and conformity assessment route.
David Small BSc (Hons), MSc, MTOPRA
Reviewed by
David Small, BSc (Hons), MSc, MTOPRA
Founder & CEO |
20+ years in medical device regulatory affairs, MDR/IVDR compliance and quality systems.
Patient Guards Recent Posts
FDA Medical Device Classification: Class I, Class II and Class III Explained
Does your medical device require FDA 510(k) clearance? This step-by-step guide explains the 510(k) process, substantial equivalence, predicate devices, testing, eSTAR submissions, FDA review and what manufacturers need to do after clearance.

FDA 510(k) Clearance: A Step-by-Step Guide for Medical Device Manufacturers
Does your medical device require FDA 510(k) clearance? This step-by-step guide explains the 510(k) process, substantial equivalence, predicate devices, testing, eSTAR submissions, FDA review and what manufacturers need to do after clearance.
FDA Medical Device Regulations: A Complete Guide for Manufacturers
Planning to place a medical device on the US market? This complete guide explains FDA medical device regulations, including device classification, 510(k), De Novo and PMA pathways, QMSR and ISO 13485, establishment registration, device listing, U.S. Agent requirements, UDI and post-market obligations.
Patient Guards Related Services
Patient Guards Regulatory Tools
Need Training?
Do you need training on Quality Management Systems or EU MDR/ EU IVDR? then check out our training courses.
 Posted on Google

 Nafiul Shelim21 days ago
Trustindex verifies that the original source of the review is Google.
I worked with Eleanor Shackleton, from Patient Guard, for the purpose of MDR, CE and UKCA marking. Her diligence, and knowledge in clinical and regulatory requirements for medical device software was critical for us. Would highly recommendPosted on Google Jay Verma46 days agoI found Patient Guard Ltd to be an exceptional partner. Their assessment was thorough, their guidance clear, and their support instrumental in helping us achieve our objectives. Steve and Ellie, in particular, were outstanding in steering us through the MHRA Class I medical device registration process.Posted on Google
 Munna P99 days agoWorking with the Patient Guard team has been a great experience throughout our MHRA and ISO 13485 documentation journey. Their expertise, structured approach, and practical guidance helped our team build a robust quality management system while keeping us aligned with regulatory expectations. The collaboration was professional, responsive, and focused on finding solutions rather than simply identifying issues. A special thank you to Alex and Steve for their outstanding coordination, responsiveness, and continuous support throughout the project. They were always approachable, provided valuable feedback, and worked closely with our team to resolve challenges efficiently. Their commitment made a significant difference in keeping our documentation effort on track. I highly recommend Patient Guard to any healthcare or MedTech organization looking for experienced regulatory and quality system partners for MHRA, ISO 13485, and broader medical device compliance initiatives. Thank you again to the entire Patient Guard team for being such reliable partners.Posted on Google
 Peter Reeve126 days agoSTEPPER design, manufacture & distribute eyewear across the globe. With the increasingly complex landscape concerning the placing of Mecial Devices onto the market, we realised we needed professional guidance. We found Patient Guard via a simple internet search and are delighted we did! They provide a pragmatic solution to our needs, are totally reliable & always available to answer our (often simplistic) questions. They are highly efficient & responsive to what is a changing picture in our world and nothing is too much trouble. We have a much better understanding of regulatory affairs and our responsibilities as manufacturers & distributors and they support us in navigating the requirements in different territories. Updating our Declaration of Conformity, ensuring our labelling is compliant and acting as our PRRC are the key areas of their service for us.Posted on Google
 Derek Timm126 days agoFor those companıes lookıng to comply to ISO standards and ın partıcular ISO13485 whıch to be honest ıs a nıghtmare I would strongly suggest goıng to the professıonals as ındeed we dıd by joınıng forces wıth Patıent Guard Ltd The staff are fantastıc nothıng ıs too much trouble and as a medıcal supply company we sımply cannot lıve wıthout them Thanks ın partıcular to Alex and Steve for all the hard work and our best regards from Dan Medıca South LımıtedPosted on Google
 BMSCriticalCare163 days agoGreat service, very helpful and always willing to answer any questions we have,Posted on Google
 Thomson Software834 days agoAlex Lewis of PatientGuard guided us through the ISO13485 process in a thorough, systematic and efficient manner. He was friendly, patient and willing to go the extra mile. Excellent service.Posted on Google
 Hannah Maddison937 days agoFantastic, knowledgeable team that are always there to help. My appointments have always been booked in very promptly and have always ended with all my queries resolved. I have found the team very flexible and their breadth of knowledge is second to none. Patient Guard are without doubt my go-to for all the regulatory aspects of my medical device role.Posted on Google
 Richard Crow972 days agoPatientguard are an excellent source of Medical regulatory compliance advice, we have taken advantage of their various services from their EU Rep service, to helping with Technical Files all the way through to using their ISO Templates to implement our ISO 13485 system.Posted on Google
 George Kitching975 days agoDavid Small and PatientGuard have been extremely helpful and supportive in assisting us with producing and updating our Technical File and Appendices for MDR certification.Verified by TrustindexTrustindex verified badge is the Universal Symbol of Trust. Only the greatest companies can get the verified badge who has a review score above 4.5, based on customer reviews over the past 12 months. Read more