Patient Guard News

Illustration showing the role of an EU Authorised Representative under the EU MDR and IVDR, depicting a non-EU manufacturer, the Authorised Representative, Article 11 compliance documentation, EUDAMED registration, Competent Authority communication and medical device regulatory oversight within the European Union.

EU Authorised Representative

Manufacturers based outside the European Union must appoint an EU Authorised Representative (EU AR or EC REP) before placing medical devices or in vitro diagnostic medical devices on the European market under the Medical Device Regulation (EU) 2017/745 (MDR) and In Vitro Diagnostic Medical Device Regulation (EU) 2017/746 (IVDR). Acting as the manufacturer’s legal representative within the EU, the Authorised Representative performs important regulatory duties, supports communication with Competent Authorities and shares legal responsibility for compliance. This guide explains who needs an EU Authorised Representative, their legal obligations under Article 11, and how to choose the right partner.

EU Authorised Representative Read More »
Regulatory workspace illustrating EU MDR Article 120 transitional provisions, featuring a transition timeline, key MDR deadlines calendar, Regulation (EU) 2017/745 open to Article 120, legacy medical device technical documentation, ISO 13485 Quality Management System files, CE-marked medical device packaging, transition compliance checklist and project plan for MDR compliance.

EU MDR Article 120 Explained: Transitional Provisions and Extended Deadlines

Article 120 of the EU Medical Device Regulation (MDR) introduced transitional provisions allowing certain legacy devices to remain on the market beyond the original MDR application date. However, these extended deadlines are only available to manufacturers that satisfy specific regulatory conditions. This guide explains how Article 120 works, which devices qualify, the current transition deadlines, what constitutes a significant change, and the practical steps manufacturers should take to maintain compliance under the EU MDR.

EU MDR Article 120 Explained: Transitional Provisions and Extended Deadlines Read More »

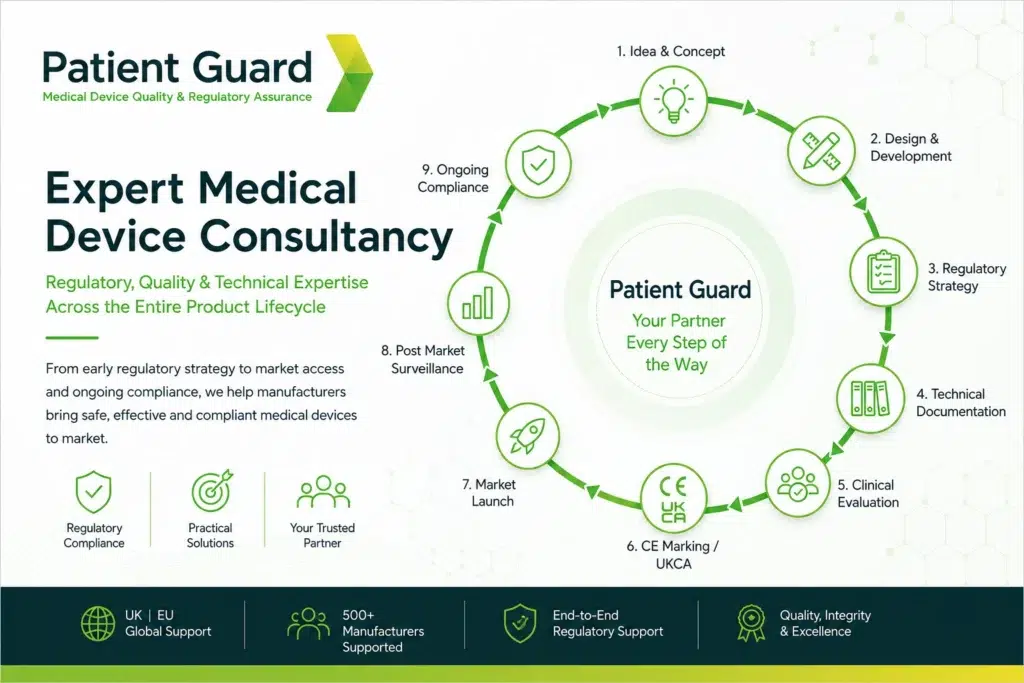
Patient Guard medical device consultancy lifecycle showing support from idea and concept through design, regulatory strategy, Technical Documentation, Clinical Evaluation, CE marking, market launch and ongoing compliance.

Medical Device Consultancy

Medical device consultancy helps manufacturers navigate complex regulatory, quality and technical requirements throughout the product lifecycle. From early regulatory strategy and product classification through to Technical Documentation, ISO 13485, clinical evaluation, biological evaluation, validation, CE marking, UKCA compliance and post-market surveillance, the right consultancy partner can reduce risk, avoid costly delays and support successful market access.

Patient Guard provides specialist medical device consultancy services for manufacturers, start-ups, distributors and international companies placing devices on the UK, EU and global markets. Our team supports organisations with practical regulatory and quality assurance guidance designed to help bring safe, compliant and effective medical devices to market.

Medical Device Consultancy Read More »

Professional reviewing medical device regulatory documentation, illustrating the role of the Person Responsible for Regulatory Compliance (PRRC) under Article 15 of EU MDR 2017/745 and IVDR 2017/746, including technical documentation, post-market surveillance and regulatory oversight.

Person Responsible for Regulatory Compliance (PRRC) under EU MDR & IVDR

The Person Responsible for Regulatory Compliance (PRRC) is a mandatory role under Article 15 of the EU Medical Device Regulation (EU MDR 2017/745) and In Vitro Diagnostic Medical Device Regulation (IVDR 2017/746). This guide explains who needs a PRRC, the required qualifications, key responsibilities, when the role can be outsourced, and how manufacturers can demonstrate compliance with regulatory requirements.

Person Responsible for Regulatory Compliance (PRRC) under EU MDR & IVDR Read More »

Get the Medical Device Technical Checklist

Thank you! The checklist is now ready to download.