EU Regulations

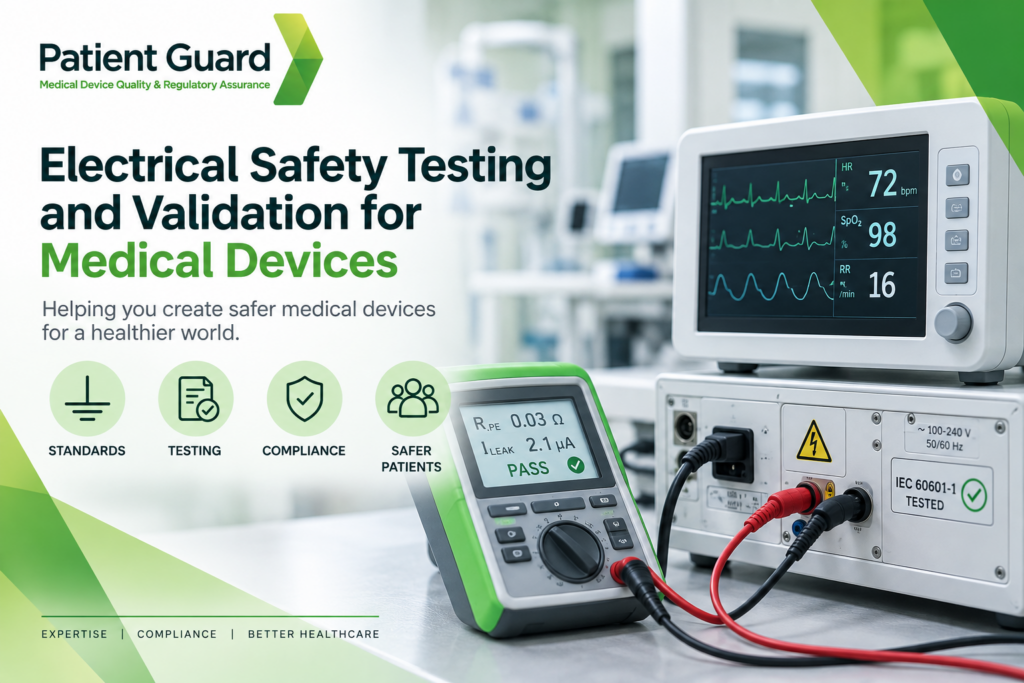
Electrical safety testing of a medical device in accordance with IEC 60601-1 requirements.

Electrical Safety Testing and Validation for Medical Devices

Electrical safety testing is essential for demonstrating the safety and performance of electrically powered medical devices. This guide explains IEC 60601-1, EMC testing, risk management, applicable standards and the evidence manufacturers need to support regulatory compliance in the UK, EU and other major markets.

Electrical Safety Testing and Validation for Medical Devices Read More »

Software as a Medical Device (SaMD)

Software as a Medical Device (SaMD): Regulatory & Development Guide

Software as a Medical Device (SaMD) has fundamentally changed the clinical landscape. From AI-driven diagnostic image scanning to standalone smartphone apps used for real-time clinical decision-making, digital health tools are scaling faster than ever. However, understanding exactly how SaMD is regulated, classified, and maintained is critical to securing market access under current MHRA and EU Notified Body expectations.

Software as a Medical Device (SaMD): Regulatory & Development Guide Read More »

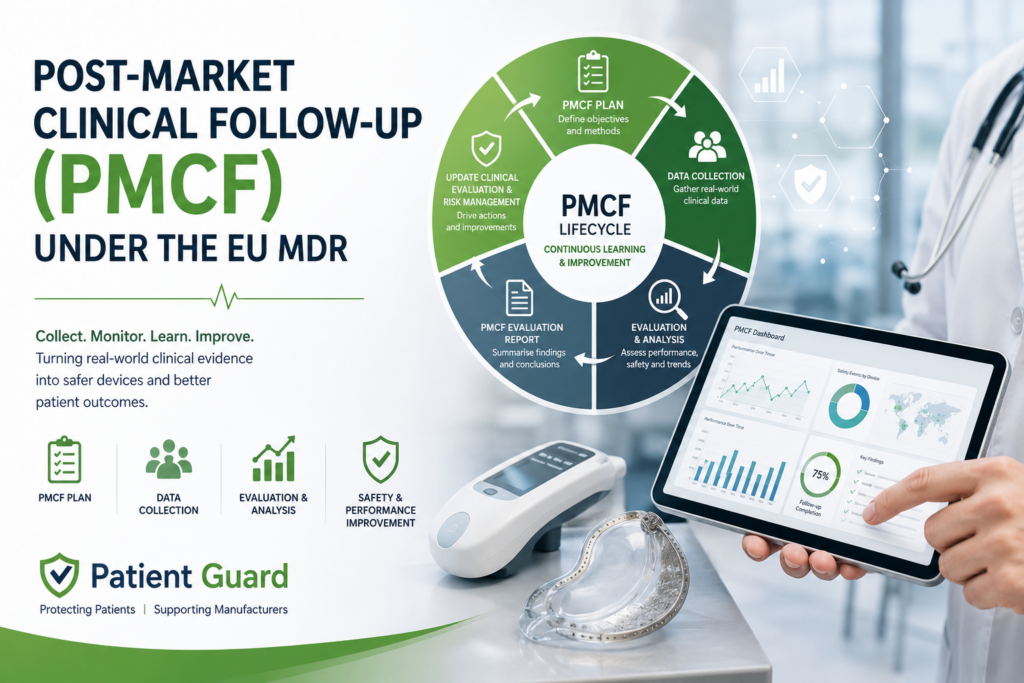
Post-Market Clinical Follow-up infographic showing the PMCF lifecycle under the EU MDR, including PMCF planning, clinical data collection, evaluation, reporting and updates to Clinical Evaluation and risk management.

Post-Market Clinical Follow-up (PMCF) Under the EU MDR

A practical guide to Post-Market Clinical Follow-up (PMCF) under the EU MDR, covering PMCF Plans, data collection methods, PMCF Evaluation Reports and how post-market clinical evidence feeds into Clinical Evaluation, risk management and ongoing medical device compliance.

Post-Market Clinical Follow-up (PMCF) Under the EU MDR Read More »

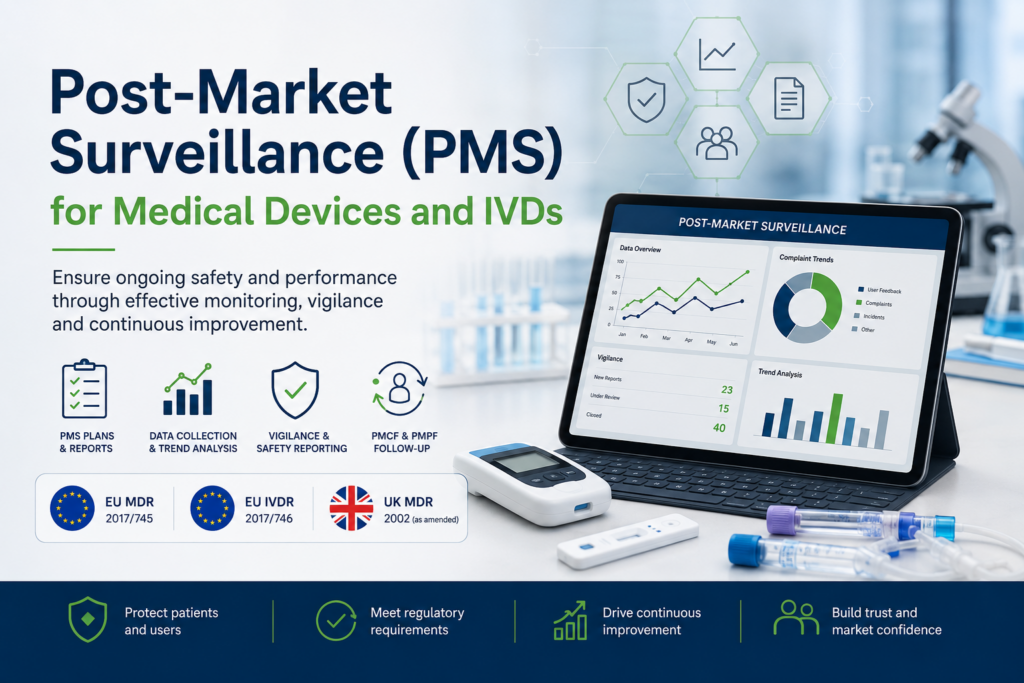
Post-market surveillance for medical devices and IVDs under the EU MDR, EU IVDR and UK MDR.

Post-Market Surveillance (PMS) for Medical Devices and IVDs

A comprehensive guide to post-market surveillance under the EU MDR, EU IVDR and UK MDR. Understand PMS plans, PMS reports, PSURs, vigilance, PMCF, PMPF and how manufacturers must monitor the safety and performance of medical devices and IVDs throughout their lifecycle.

Post-Market Surveillance (PMS) for Medical Devices and IVDs Read More »

Illustration showing the medical device clinical evaluation process under the EU MDR, with scientific literature, clinical investigations, Post-Market Surveillance, Post-Market Clinical Follow-up, risk management and Clinical Evaluation Reports combining to demonstrate device safety, clinical performance and an acceptable benefit-risk profile for CE marking.

Medical Device Clinical Evaluation: A Complete Guide

Clinical Evaluation is a continuous process used to demonstrate that a medical device is safe, achieves its intended clinical performance and maintains an acceptable benefit-risk profile throughout its lifecycle. This complete guide explains every stage of clinical evaluation under the EU MDR, including Clinical Evaluation Plans (CEPs), Clinical Evaluation Reports (CERs), clinical evidence, literature reviews, equivalence, PMCF, PMS and ongoing regulatory compliance.

Medical Device Clinical Evaluation: A Complete Guide Read More »

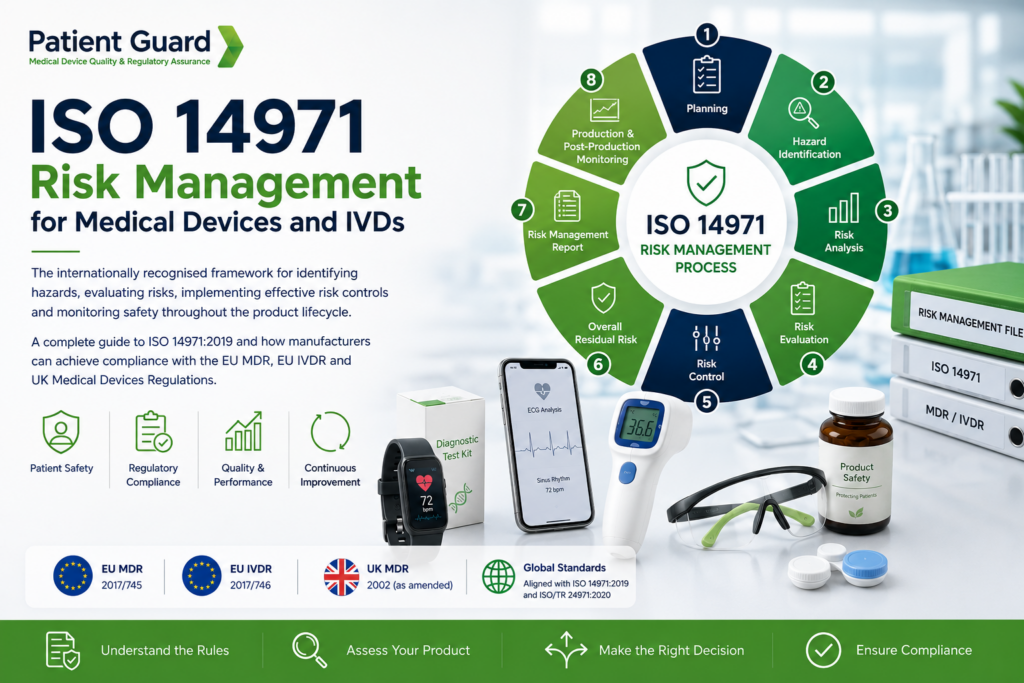
Professional hero banner illustrating the ISO 14971 risk management process for medical devices and in vitro diagnostic medical devices, showing the eight stages of risk management including planning, hazard identification, risk analysis, risk evaluation, risk control, residual risk evaluation, risk management reporting and post-production monitoring to support compliance with the EU MDR, EU IVDR and UK Medical Devices Regulations.

ISO 14971 Risk Management for Medical Devices and IVDs

ISO 14971 provides the internationally recognised framework for identifying hazards, evaluating risks, implementing effective risk controls and monitoring safety throughout the lifecycle of a medical device or in vitro diagnostic device.

This comprehensive guide explains how manufacturers can apply ISO 14971:2019 in practice, maintain a compliant Risk Management File and demonstrate conformity with the EU MDR, EU IVDR and UK Medical Devices Regulations.

ISO 14971 Risk Management for Medical Devices and IVDs Read More »

Illustration showing the role of an EU Authorised Representative under the EU MDR and IVDR, depicting a non-EU manufacturer, the Authorised Representative, Article 11 compliance documentation, EUDAMED registration, Competent Authority communication and medical device regulatory oversight within the European Union.

EU Authorised Representative

Manufacturers based outside the European Union must appoint an EU Authorised Representative (EU AR or EC REP) before placing medical devices or in vitro diagnostic medical devices on the European market under the Medical Device Regulation (EU) 2017/745 (MDR) and In Vitro Diagnostic Medical Device Regulation (EU) 2017/746 (IVDR). Acting as the manufacturer’s legal representative within the EU, the Authorised Representative performs important regulatory duties, supports communication with Competent Authorities and shares legal responsibility for compliance. This guide explains who needs an EU Authorised Representative, their legal obligations under Article 11, and how to choose the right partner.

EU Authorised Representative Read More »

Illustration showing the biological evaluation of a medical device under ISO 10993, highlighting the risk-based process from the Biological Evaluation Plan (BEP) through biological assessment, Biological Evaluation Report (BER), Post-Market Surveillance (PMS), and integration with ISO 14971 Risk Management throughout the medical device lifecycle.

Biological Evaluation of Medical Devices: Complete ISO 10993 Guide

Biological Evaluation is a risk-based process used to demonstrate that a medical device is biologically safe for its intended clinical use. This complete guide explains how ISO 10993 supports Biological Evaluation, when biocompatibility testing is required, how Biological Evaluation Plans (BEPs) and Biological Evaluation Reports (BERs) are developed, and how Biological Evaluation integrates with Risk Management and Clinical Evaluation throughout the device lifecycle.

Biological Evaluation of Medical Devices: Complete ISO 10993 Guide Read More »

Get the Medical Device Technical Checklist

Thank you! The checklist is now ready to download.