EU Regulations

Professional illustration of the CE marking process for medical devices and in vitro diagnostic medical devices, showing technical documentation, risk management, clinical evaluation, quality management, an EU Declaration of Conformity and a CE marked medical device under the EU MDR 2017/745 and IVDR 2017/746.

CE Marking Medical Devices: A Complete Guide to EU MDR & IVDR Compliance

CE marking demonstrates that a medical device or in vitro diagnostic medical device complies with the applicable requirements of the EU Medical Device Regulation (EU MDR 2017/745) or In Vitro Diagnostic Medical Device Regulation (IVDR 2017/746). This guide explains the CE marking process, device classification, conformity assessment, technical documentation, clinical and performance evaluation, Notified Body involvement, post-market surveillance and the steps manufacturers must complete before placing a device on the European market.

CE Marking Medical Devices: A Complete Guide to EU MDR & IVDR Compliance Read More »

Hero image illustrating the process of determining whether a product is a medical device, featuring examples of medical and non-medical products alongside CE Marking and UKCA symbols, highlighting intended purpose, mode of action, medical claims, borderline products and the regulatory pathway under the EU MDR and UK Medical Devices Regulations.

Is My Product a Medical Device? How to Determine Whether the EU MDR or UK Regulations Apply

Determining whether your product is a medical device is the first and most important step towards regulatory compliance. This guide explains how regulators assess intended purpose, mode of action and medical claims, helping you understand whether your product falls within the scope of the EU Medical Device Regulation (MDR), the UK Medical Devices Regulations or the IVDR. Whether you’re developing software, wearable technology, wellness products, cosmetics or innovative healthcare solutions, this article will help you identify the correct regulatory pathway and avoid common classification mistakes.

Is My Product a Medical Device? How to Determine Whether the EU MDR or UK Regulations Apply Read More »

Skin Irritation Testing in the biological evaluation of medical devices (ISO 10993-23)

ISO 10993-23 Skin Irritation Testing Explained: Methods, Requirements and Regulatory Expectations

Skin irritation occurs when a substance causes inflammation or damage to the skin upon contact. For medical devices, this can be a serious concern, as they are often in prolonged contact with the skin. The consequences of not adequately testing for irritation can range from minor redness to severe dermatitis, leading to discomfort, injury, or even infection in users. Therefore, reliable testing methods are crucial to ensure that products are safe before they reach the market.

ISO 10993-23 Skin Irritation Testing Explained: Methods, Requirements and Regulatory Expectations Read More »

Cytotoxicity Testing in Medial Devices ISO 10993-5

Cytotoxicity Testing in Medical Device Biological Evaluation

In the highly regulated world of medical devices, ensuring patient safety is paramount. One of the essential components of this safety assurance is cytotoxicity testing, specifically adhering to ISO 10993-5 standards. This blog explores the importance of ISO 10993-5 cytotoxicity testing, particularly for medical devices that come into direct contact with patients, and how it contributes to the overall safety and efficacy of these devices.

Cytotoxicity Testing in Medical Device Biological Evaluation Read More »

drug Infusion bag

Drug Stability in Medical Devices: Syringes, Infusion Sets and Bags

Ensuring the stability of drugs in contact with medical devices such as syringes, infusion administration sets and infusion administration bags is paramount. This article delves into the importance of drug stability testing, the challenges faced, and the methodologies employed to guarantee that the drugs administered through these devices remain safe and effective.

Drug Stability in Medical Devices: Syringes, Infusion Sets and Bags Read More »

E&L Testing in medical devices

ISO 10993 Extractables and Leachables Testing: Chemical Characterisation and Toxicological Risk Assessment

In medical device design and development and manufacturing, ensuring the safety and biocompatibility of products is paramount. As part of the biological evaluation process, extractables and leachables (E&L) testing plays a crucial role. This testing ensures that any chemicals that might migrate from the device materials into the body do not pose a risk to patients. The ISO 10993 standard series provides comprehensive guidelines for the biological evaluation of medical devices, including protocols for E&L testing. In this blog, we will delve into the importance of extractables and leachables testing, its procedures, and its relevance within the ISO 10993 framework.

ISO 10993 Extractables and Leachables Testing: Chemical Characterisation and Toxicological Risk Assessment Read More »

Regulatory workspace illustrating EU MDR Article 120 transitional provisions, featuring a transition timeline, key MDR deadlines calendar, Regulation (EU) 2017/745 open to Article 120, legacy medical device technical documentation, ISO 13485 Quality Management System files, CE-marked medical device packaging, transition compliance checklist and project plan for MDR compliance.

EU MDR Article 120 Explained: Transitional Provisions and Extended Deadlines

Article 120 of the EU Medical Device Regulation (MDR) introduced transitional provisions allowing certain legacy devices to remain on the market beyond the original MDR application date. However, these extended deadlines are only available to manufacturers that satisfy specific regulatory conditions. This guide explains how Article 120 works, which devices qualify, the current transition deadlines, what constitutes a significant change, and the practical steps manufacturers should take to maintain compliance under the EU MDR.

EU MDR Article 120 Explained: Transitional Provisions and Extended Deadlines Read More »

A certificate that says 'ISO 13485 certified' on it - This image is used by patient guard to represent ISO 13485 quality assurance and QMS services.

What is ISO 13485? The Complete Guide to Medical Device QMS

ISO 13485:2016 is the international standout standard that defines the precise requirements for a Quality Management System (QMS) specific to the medical device industry. Unlike general business quality frameworks, it prioritizes patient safety, clinical efficacy, and global regulatory compliance ahead of standard consumer satisfaction metrics.

Whether you are an active medical device manufacturer, a software-as-a-service (SaMD) developer, an importer, or an EU/UK distributor, demonstrating compliance with this standard is virtually mandatory to secure and retain critical market access.

What is ISO 13485? The Complete Guide to Medical Device QMS Read More »

Infographic illustrating the different medical device registration pathways for the European Union, United Kingdom and international markets, showing the relationship between CE marking, EUDAMED registration, UDI assignment, MHRA registration, UK Responsible Person, Authorised Representative and international regulatory registrations including FDA and ARTG.

Medical Device Registration Explained: EU, UK and Global Requirements

Medical device registration is not a single regulatory process. Depending on where you intend to market your medical device or IVD, you may need CE marking, MHRA registration, EUDAMED registration, UDI assignment or an Authorised Representative. This guide explains the different registration requirements under the EU MDR, IVDR and UK regulations, helping manufacturers understand which obligations apply before placing their products on the market.

Medical Device Registration Explained: EU, UK and Global Requirements Read More »

Hero image illustrating the structure of a medical device technical file under the EU MDR, showing the key documentation required for CE marking, including device description, General Safety and Performance Requirements (GSPRs), Risk Management File, verification and validation, Clinical Evaluation, Biological Evaluation and Post-Market Surveillance in accordance with Regulation (EU) 2017/745.

How to Structure a Medical Device Technical File

A medical device technical file is a concise set of documents created by a manufacturer to explain the performance and safety of a particular Medical Device in a clear, well-organized, easily searchable, and unambiguous manner.

In order to prove compliance with the general safety and performance criteria of the legislation, the manufacturer must have – and keep up-to-date – this technical documentation, regardless of  medical device class.

How to Structure a Medical Device Technical File Read More »

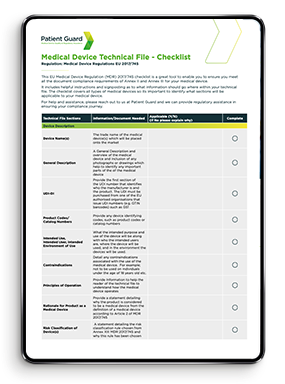
Get the Medical Device Technical Checklist

Thank you! The checklist is now ready to download.