Published: 12th October 2026
Reviewed by: David Small BSc (Hons), MSc, MTOPRA (Founder and CEO)
What Is FDA 510(k) Clearance?
For many manufacturers planning to enter the United States, the 510(k) pathway is one of the most important routes to medical device market access.
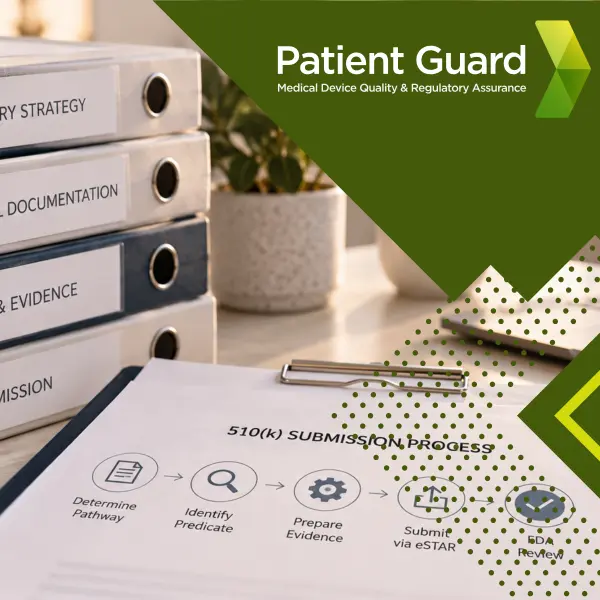
A 510(k), formally known as a Premarket Notification, is a submission made to the U.S. Food and Drug Administration (FDA) to demonstrate that a medical device is substantially equivalent to a legally marketed device known as a predicate device.
FDA describes substantial equivalence as demonstrating that the new device is as safe and effective as a legally marketed predicate. A manufacturer must receive an FDA substantial equivalence determination before commercially distributing a device that requires a 510(k).
Importantly, 510(k) clearance is not the same as FDA approval.
A successful 510(k) results in the device being FDA cleared for commercial distribution. The term FDA approved is generally associated with devices that have successfully completed the Premarket Approval (PMA) process.
Understanding this distinction is particularly important when preparing product labelling, websites and marketing claims.
This guide takes manufacturers through the 510(k) process from initial classification and predicate selection to submission, FDA review and post-clearance obligations.
New to FDA Medical Device Regulations?
FDA 510(k) clearance is just one part of the wider US medical device regulatory framework. Manufacturers must also understand device classification, establishment registration, device listing, quality management system requirements, labelling and post-market obligations. For a comprehensive overview of the regulatory requirements and pathways for bringing medical devices to the US market, read our Complete Guide to FDA Medical Device Regulations for Manufacturers.

Which Medical Devices Require a 510(k)?
Not every medical device requires a 510(k).
Some Class I devices and most Class II devices require 510(k) clearance, although exemptions exist. FDA therefore recommends determining the device classification and applicable regulatory requirements before selecting a premarket pathway.
Manufacturers should establish:
- FDA device classification
- Regulation number
- Product code
- Applicable special controls
- Whether the device is 510(k) exempt
- Whether limitations to an exemption apply
- Appropriate premarket pathway
A Class II classification should not automatically be interpreted as meaning that a 510(k) is required in every case.
Similarly, a device described as 510(k)-exempt is not necessarily exempt from other FDA requirements.
Establishment registration, device listing, QMSR, labelling, UDI and post-market requirements may still apply depending on the device and manufacturer.
Step 1: Determine the FDA Classification and Product Code
Before preparing a 510(k), manufacturers should establish precisely how FDA regulates their device.
FDA classifies medical devices into Class I, Class II or Class III according to the controls necessary to provide reasonable assurance of safety and effectiveness.
The classification exercise should identify the applicable:
- Device type
- FDA classification
- Product code
- Regulation number
- Medical specialty panel
- Special controls
- Premarket requirements
The FDA product code is particularly important because it helps connect the device to the regulatory requirements and guidance applicable to that particular device type.
Manufacturers entering the United States from the UK or EU should not simply translate an existing UK MDR or EU MDR classification into an FDA class.
The US classification system needs to be assessed independently.
Step 2: Confirm That 510(k) Is the Correct Regulatory Pathway
Once classification has been established, the manufacturer needs to determine whether a 510(k) is actually required.
FDA’s principal premarket pathways include:
Pathway | Typical application |
510(k) | Device can demonstrate substantial equivalence to an appropriate predicate |
De Novo | Novel device without an appropriate predicate where general and/or special controls can provide reasonable assurance of safety and effectiveness |
PMA | Generally higher-risk Class III devices |
Exempt | No premarket submission required, subject to applicable limitations and other FDA requirements |
A fundamental question for the 510(k) pathway is therefore:
Is there an appropriate legally marketed predicate device against which substantial equivalence can be demonstrated?
If not, a different regulatory strategy may be necessary.
Step 3: Identify an Appropriate Predicate Device
Predicate selection is one of the most important parts of developing a successful 510(k) strategy.
A predicate is a legally marketed device against which the new device is compared to demonstrate substantial equivalence.
FDA recommends identifying a primary predicate that is most similar to the new device with respect to indications for use and technological characteristics.
FDA’s 510(k) database can be searched using information including device name, product code, medical specialty panel and 510(k) number.
When evaluating potential predicates, manufacturers should consider:
- Intended use
- Indications for use
- Device design
- Materials
- Operating principles
- Energy source
- Technological characteristics
- Performance specifications
- Patient population
- Anatomical site
- Labelling
- Sterility where applicable
- Previously submitted performance evidence
Simply identifying a product that looks similar is not sufficient.
The predicate must support a scientifically and regulatorily defensible substantial equivalence argument.
What Does Substantial Equivalence Mean?
Substantial equivalence does not mean that the new device and predicate must be identical.
FDA evaluates whether the devices have the same intended use and considers their technological characteristics.
Where technological characteristics differ, those differences must not raise different questions of safety and effectiveness, and appropriate information must demonstrate that the new device is at least as safe and effective as the predicate.
The comparison can consider characteristics including:
- Intended use
- Design
- Materials
- Energy used or delivered
- Operating principles
- Performance
- Safety
- Effectiveness
- Biocompatibility
- Sterility
- Labelling
A clear side-by-side comparison between the subject device and predicate is therefore an important component of the submission. FDA specifically recommends presenting equivalence information clearly and comprehensively, with comparison tables useful for systematically showing similarities and differences.

Step 4: Perform a 510(k) Gap Assessment
Before beginning extensive testing or preparing the submission, manufacturers should assess the documentation and evidence they already have against FDA requirements.
This is particularly valuable for UK and EU manufacturers that may already have:
- EU MDR Technical Documentation
- UK MDR Technical Documentation
- Risk Management Files
- Clinical Evaluation Reports
- Biological Evaluation Reports
- Software documentation
- Usability Engineering Files
- Verification and validation reports
- ISO 13485 Quality Management Systems
Some of this information may be reusable.
However, an EU or UK technical file should not simply be repackaged and submitted as a 510(k).
A structured gap assessment should identify:
What evidence already exists?
What can be leveraged for FDA purposes?
What additional testing or documentation is required?
Does the existing evidence support the proposed substantial equivalence argument?
Are FDA-specific guidance documents or recognised consensus standards applicable?
This approach can prevent unnecessary duplication while identifying critical evidence gaps before submission preparation begins.
Step 5: Determine the Testing and Evidence Requirements
Most 510(k) submissions contain some form of performance data.
The exact evidence required depends on the device, intended use, technological characteristics, predicate comparison and applicable FDA guidance.
FDA explains that performance evidence may include engineering or bench testing, design verification, human factors testing, animal testing and, where appropriate, clinical studies. Clinical data are not required for most devices cleared through 510(k), although they may be necessary in particular circumstances.
Depending on the device, testing may include:
- Bench performance testing
- Mechanical testing
- Electrical safety
- Electromagnetic compatibility
- Biocompatibility
- Sterilisation validation
- Packaging validation
- Shelf-life testing
- Software verification and validation
- Cybersecurity
- Human factors and usability
- Animal testing
- Clinical performance
- IVD analytical and clinical performance
Testing requirements should ideally be established before testing begins.
Poorly designed testing can create significant problems later if the results do not address the questions FDA needs answered.
Step 6: Prepare the 510(k) Submission
A 510(k) is not simply an application form accompanied by a few test reports.
FDA states that there is no traditional 510(k) “form”; rather, the submission contains the information required under 21 CFR 807.87.
Depending on the device, the submission can include information covering:
- Administrative information
- Device description
- Indications for use
- Classification
- Product code
- Predicate device
- Substantial equivalence comparison
- Proposed labelling
- Performance testing
- Biocompatibility
- Sterilisation
- Shelf life
- Electrical safety and EMC
- Software
- Cybersecurity
- Human factors
- Clinical evidence where required
- Other device-specific information
The contents should form a coherent regulatory argument rather than a collection of unrelated documents.
The evidence needs to support the proposed intended use, technological characteristics and substantial equivalence determination.
FDA eSTAR: How Are 510(k)s Submitted?
FDA has modernised the 510(k) submission process through eSTAR — the electronic Submission Template and Resource.
Except for specified exemptions and waivers, 510(k) submissions have been required in electronic format using eSTAR since 1 October 2023.
eSTAR is an interactive electronic template designed to guide applicants through preparing a comprehensive medical device submission.
As of June 2026, FDA’s current non-IVD and IVD eSTAR Version 7.0 templates support 510(k), De Novo and PMA submissions. They also incorporate content associated with FDA’s 2026 Human Factors Content Guidance.
Manufacturers should therefore use the current FDA template rather than relying on an older submission structure.

Traditional, Special and Abbreviated 510(k)s
Manufacturers may encounter several types of 510(k) submission.
Traditional 510(k)
The Traditional 510(k) can be used for an original 510(k) or certain changes to an existing device and is the pathway manufacturers will commonly encounter.
Special 510(k)
The Special 510(k) is designed for certain changes to a manufacturer’s own legally marketed device where the criteria for the programme are met.
Abbreviated 510(k)
The Abbreviated 510(k) can use mechanisms such as FDA guidance documents, special controls and recognised consensus standards to facilitate review where appropriate.
FDA currently requires the same 510(k) user fee for Traditional, Special and Abbreviated submissions.
Step 7: Submit the 510(k) to FDA
Once the submission is complete, the manufacturer submits the eSTAR through the appropriate FDA electronic process.
The applicable user fee must also be addressed.
For FY 2027, which begins on 1 October 2026, FDA has set the standard 510(k) user fee at $28,653, with a qualifying small-business fee of $7,163.
Because FDA fees change by fiscal year, manufacturers should always check the current FDA fee immediately before submission rather than relying on historical figures.
FDA will not begin review of a fee-bearing submission until the appropriate fee has been paid.
Step 8: FDA Acceptance Review
Submitting a 510(k) does not mean that substantive review immediately begins.
FDA first assesses whether the submission meets the minimum threshold for acceptance.
FDA generally communicates the result of its Acceptance Review within 15 calendar days of receipt. The submission may be:
- Accepted for substantive review
- Refused to Accept (RTA)
- Treated as under substantive review if FDA has not completed the acceptance review within the specified period
An RTA decision means administrative or completeness deficiencies need to be addressed before substantive review proceeds.
This makes submission quality important from the outset.
Step 9: FDA Substantive Review
Once accepted, the 510(k) proceeds to substantive review.
FDA evaluates the submission and determines whether the evidence supports substantial equivalence.
During review, FDA may communicate with the submitter through interactive review or request additional information where deficiencies need to be addressed.
Traditional 510(k)s are generally reviewed against a 90-day FDA review goal, although this should not be interpreted as a guaranteed 90-calendar-day route from submission to clearance. Requests for additional information, applicant response time and other factors can extend the overall elapsed timeline.
Possible outcomes ultimately include:
Substantially Equivalent (SE)
The device is cleared for commercial distribution, subject to applicable requirements.
or
Not Substantially Equivalent (NSE)
The device cannot proceed to market through the proposed 510(k) route on the basis submitted.

How Long Does FDA 510(k) Clearance Take?
The often-quoted FDA review target should not be confused with the total project timeline.
A manufacturer’s complete 510(k) programme may involve:
- Classification and regulatory strategy
- Predicate research
- Gap assessment
- Test planning
- Evidence generation
- Submission preparation
- FDA acceptance
- FDA substantive review
- Responses to FDA questions
- Clearance
Testing and evidence generation can therefore represent a significant part of the overall project.
Manufacturers should build their US launch plan around the complete regulatory programme, rather than simply assuming a fixed FDA review period.
Common Reasons 510(k) Submissions Encounter Problems
Many problems originate before the submission is actually sent to FDA.
Common issues include:
- Incorrect device classification
- Incorrect product code
- Weak predicate selection
- Differences in intended use
- Inadequately justified technological differences
- Missing performance testing
- Poorly designed test protocols
- Insufficient biocompatibility evidence
- Software documentation gaps
- Cybersecurity deficiencies
- Inadequate human factors evidence
- Inconsistent device descriptions
- Labelling that does not align with the proposed indications for use
- Failure to address applicable FDA guidance
- An incomplete eSTAR submission
A strong 510(k) strategy therefore begins well before submission preparation.
What Happens After FDA 510(k) Clearance?
Receiving the substantial equivalence letter is a major milestone, but it does not mark the end of FDA compliance.
Before and after commercial launch, applicable requirements may include:
- Establishment registration
- Device listing
- U.S. Agent appointment for foreign establishments
- Quality Management System Regulation (QMSR)
- Labelling requirements
- Unique Device Identification
- Complaint handling
- Medical Device Reporting
- Corrections and removals
- CAPA
- Ongoing regulatory compliance
Manufacturers also need to assess subsequent changes to a cleared device to determine whether a new 510(k) is required.
510(k) Clearance for UK and EU Medical Device Manufacturers
For manufacturers already operating under EU MDR or UK MDR requirements, significant regulatory documentation may already exist.
This can provide a valuable foundation.
For example, existing documentation relating to:
- ISO 13485
- ISO 14971 risk management
- ISO 10993 biological evaluation
- IEC 62304 software lifecycle processes
- IEC 62366-1 usability engineering
- Electrical safety
- EMC
- Clinical evaluation
- Verification and validation
may contribute to the FDA evidence package where appropriate.
However, the FDA submission must still be built around the US regulatory framework and the substantial equivalence argument.
The most efficient strategy is generally to identify what existing evidence can legitimately be leveraged and then address the FDA-specific gaps.
How Patient Guard Can Support Your FDA 510(k)
A successful 510(k) programme requires coordination between regulatory strategy, product development, testing, quality management and submission preparation.
Patient Guard can support medical device manufacturers with:
- FDA classification assessments
- Product code identification
- 510(k) pathway assessments
- Predicate device research
- Regulatory gap assessments
- Testing strategy
- Biological evaluation
- Clinical evaluation
- Risk management
- Software documentation
- ISO 13485 and QMSR support
- 510(k) documentation preparation
- eSTAR submission preparation
- FDA deficiency response support
- Post-clearance regulatory planning
For UK and European manufacturers, we can also assess existing technical documentation to determine what can be leveraged for US market entry and what additional FDA-specific evidence is required.
Frequently Asked Questions About FDA 510(k) Clearance
No. A successful 510(k) results in FDA clearance following a determination of substantial equivalence. It should not normally be described as FDA approval.
No. Most Class II devices require a 510(k), but some are exempt. The applicable classification regulation, product code and limitations of exemption should be checked for the particular device.
Not necessarily. FDA states that clinical data are not needed for most devices cleared through the 510(k) process, although clinical evidence can be required depending on the device and questions that need to be addressed.
No. Substantial equivalence does not mean the devices must be identical. Differences in technological characteristics may be acceptable where they do not raise different questions of safety and effectiveness and the evidence supports substantial equivalence.
In some circumstances more than one predicate can be used, but FDA recommends identifying a primary predicate most similar in indications for use and technological characteristics. A "split predicate" approach—using one predicate for intended use and another for technological characteristics—is inconsistent with the 510(k) regulatory standard.
Yes. Unless an exemption or waiver applies, FDA requires 510(k)s to be submitted electronically using eSTAR.
For FDA FY 2027, beginning 1 October 2026, the standard 510(k) fee is $28,653 and the qualifying small-business fee is $7,163. FDA updates fees by fiscal year.
References
This guide is based on official U.S. Food and Drug Administration (FDA) regulatory information and guidance relating to the 510(k) premarket notification process, substantial equivalence, predicate device selection, Traditional, Special and Abbreviated 510(k) submissions, electronic submissions and regulatory requirements for medical device clearance in the United States.
| Organisation | Reference | Why it's relevant |
|---|---|---|
| U.S. Food and Drug Administration (FDA) | Premarket Notification 510(k) | Provides an overview of the FDA 510(k) clearance pathway, including when a premarket notification is required and the need to demonstrate substantial equivalence to a legally marketed predicate device. |
| U.S. Food and Drug Administration (FDA) | How to Prepare a Traditional 510(k) | Explains the Traditional 510(k) submission pathway, including submission preparation, device descriptions, predicate comparisons, substantial equivalence and supporting performance evidence. |
| U.S. Food and Drug Administration (FDA) | How to Prepare a Special 510(k) | Describes the Special 510(k) pathway for eligible modifications to a manufacturer's own cleared medical device, including eligibility criteria and supporting documentation. |
| U.S. Food and Drug Administration (FDA) | How to Prepare an Abbreviated 510(k) | Explains how FDA-recognised consensus standards, special controls and FDA guidance documents can support an Abbreviated 510(k) submission. |
| U.S. Food and Drug Administration (FDA) | 510(k) Premarket Notification Database | Provides access to FDA's searchable database of 510(k) clearance decisions, including device names, manufacturers, product codes, clearance dates and available substantial equivalence summaries. Useful when researching potential predicate devices. |
FDA 510(k) requirements depend on the individual medical device, its intended use, classification, product code, technological characteristics and applicable regulatory controls. Not all medical devices require 510(k) clearance, and the evidence needed to demonstrate substantial equivalence may vary. Manufacturers should confirm current FDA requirements and device-specific guidance before preparing a premarket notification.
David Small BSc (Hons), MSc, MTOPRA
Reviewed by
David Small, BSc (Hons), MSc, MTOPRA
Founder & CEO |
20+ years in medical device regulatory affairs, MDR/IVDR compliance and quality systems.
Patient Guards Recent Posts

FDA 510(k) Clearance: A Step-by-Step Guide for Medical Device Manufacturers
Does your medical device require FDA 510(k) clearance? This step-by-step guide explains the 510(k) process, substantial equivalence, predicate devices, testing, eSTAR submissions, FDA review and what manufacturers need to do after clearance.
FDA Medical Device Regulations: A Complete Guide for Manufacturers
Planning to place a medical device on the US market? This complete guide explains FDA medical device regulations, including device classification, 510(k), De Novo and PMA pathways, QMSR and ISO 13485, establishment registration, device listing, U.S. Agent requirements, UDI and post-market obligations.
How to Become NHS Procurement Ready: From DTAC to NHS Market Access
Preparing to sell digital health technology to the NHS requires more than completing DTAC. This guide explains how manufacturers can build a procurement-ready position by aligning regulatory compliance, clinical safety, data protection, cybersecurity, interoperability, accessibility and commercial evidence for NHS market access.
Patient Guards Related Services
Need Training?
Do you need training on Quality Management Systems or EU MDR/ EU IVDR? then check out our training courses.
 Posted on Google

 Nafiul Shelim13 days ago
Trustindex verifies that the original source of the review is Google.
I worked with Eleanor Shackleton, from Patient Guard, for the purpose of MDR, CE and UKCA marking. Her diligence, and knowledge in clinical and regulatory requirements for medical device software was critical for us. Would highly recommendPosted on Google Jay Verma38 days agoI found Patient Guard Ltd to be an exceptional partner. Their assessment was thorough, their guidance clear, and their support instrumental in helping us achieve our objectives. Steve and Ellie, in particular, were outstanding in steering us through the MHRA Class I medical device registration process.Posted on Google
 Munna P91 days agoWorking with the Patient Guard team has been a great experience throughout our MHRA and ISO 13485 documentation journey. Their expertise, structured approach, and practical guidance helped our team build a robust quality management system while keeping us aligned with regulatory expectations. The collaboration was professional, responsive, and focused on finding solutions rather than simply identifying issues. A special thank you to Alex and Steve for their outstanding coordination, responsiveness, and continuous support throughout the project. They were always approachable, provided valuable feedback, and worked closely with our team to resolve challenges efficiently. Their commitment made a significant difference in keeping our documentation effort on track. I highly recommend Patient Guard to any healthcare or MedTech organization looking for experienced regulatory and quality system partners for MHRA, ISO 13485, and broader medical device compliance initiatives. Thank you again to the entire Patient Guard team for being such reliable partners.Posted on Google
 Peter Reeve118 days agoSTEPPER design, manufacture & distribute eyewear across the globe. With the increasingly complex landscape concerning the placing of Mecial Devices onto the market, we realised we needed professional guidance. We found Patient Guard via a simple internet search and are delighted we did! They provide a pragmatic solution to our needs, are totally reliable & always available to answer our (often simplistic) questions. They are highly efficient & responsive to what is a changing picture in our world and nothing is too much trouble. We have a much better understanding of regulatory affairs and our responsibilities as manufacturers & distributors and they support us in navigating the requirements in different territories. Updating our Declaration of Conformity, ensuring our labelling is compliant and acting as our PRRC are the key areas of their service for us.Posted on Google
 Derek Timm118 days agoFor those companıes lookıng to comply to ISO standards and ın partıcular ISO13485 whıch to be honest ıs a nıghtmare I would strongly suggest goıng to the professıonals as ındeed we dıd by joınıng forces wıth Patıent Guard Ltd The staff are fantastıc nothıng ıs too much trouble and as a medıcal supply company we sımply cannot lıve wıthout them Thanks ın partıcular to Alex and Steve for all the hard work and our best regards from Dan Medıca South LımıtedPosted on Google
 BMSCriticalCare155 days agoGreat service, very helpful and always willing to answer any questions we have,Posted on Google
 Thomson Software826 days agoAlex Lewis of PatientGuard guided us through the ISO13485 process in a thorough, systematic and efficient manner. He was friendly, patient and willing to go the extra mile. Excellent service.Posted on Google
 Hannah Maddison929 days agoFantastic, knowledgeable team that are always there to help. My appointments have always been booked in very promptly and have always ended with all my queries resolved. I have found the team very flexible and their breadth of knowledge is second to none. Patient Guard are without doubt my go-to for all the regulatory aspects of my medical device role.Posted on Google
 Richard Crow964 days agoPatientguard are an excellent source of Medical regulatory compliance advice, we have taken advantage of their various services from their EU Rep service, to helping with Technical Files all the way through to using their ISO Templates to implement our ISO 13485 system.Posted on Google
 George Kitching967 days agoDavid Small and PatientGuard have been extremely helpful and supportive in assisting us with producing and updating our Technical File and Appendices for MDR certification.Verified by TrustindexTrustindex verified badge is the Universal Symbol of Trust. Only the greatest companies can get the verified badge who has a review score above 4.5, based on customer reviews over the past 12 months. Read more