Guides and Advice

ISO 9001 vs ISO 13485

ISO 9001 vs ISO 13485: Key Differences Explained

While both standards share the same core structural DNA, they serve entirely different strategic purposes. ISO 9001 is the universal language of operational quality management; ISO 13485 is its highly specialized medical-device dialect—one that speaks directly to global regulators, notified bodies, and clinical auditors.

As the European MDR/IVDR timelines advance and the US FDA’s new Quality Management System Regulation (QMSR) takes full effect, global regulatory scrutiny has intensified. Every regulatory audit now explicitly tests whether a manufacturer’s medical-device quality management system (QMS) truly mitigates risk or merely exists on paper as passive compliance documentation.

ISO 9001 vs ISO 13485: Key Differences Explained Read More »
Mastering ISO 13485 Compliance with a Lean QMS

Mastering ISO 13485 Compliance with a Lean QMS

In today’s medical-device sector, ISO 13485 compliance is no longer optional – it’s the ticket to market access, regulatory trust, and operational maturity.
As the European MDR and the US FDA’s new Quality Management System Regulation (QMSR) converge, scrutiny has intensified. Every audit now tests whether a company’s medical-device quality management system (QMS) truly works or merely exists on paper.

Mastering ISO 13485 Compliance with a Lean QMS Read More »

UDI and EUDAMED: Strengthen Medical Device Traceability

UDI and EUDAMED: Strengthen Medical Device Traceability

The alphabet soup of medical device regulation – UDI, EUDAMED, MDR – can make even the most seasoned regulatory manager twitch. But there’s no escaping it: UDI and EUDAMED are now at the heart of European medical device traceability.

UDI and EUDAMED: Strengthen Medical Device Traceability Read More »

ISO 13485 audit readiness

ISO 13485 Audit Readiness: How to Pass with Confidence

With ISO 13485 audits tightening worldwide, unprepared medical-device companies face major risks in 2026 and beyond. The regulatory climate has changed fast: global authorities are aligning standards, increasing unannounced inspections, and raising expectations of documentation quality.

ISO 13485 Audit Readiness: How to Pass with Confidence Read More »

V-58.-How-Changes-in-the-EU-MDR-Impact-Your-Supply-Chain-Agreements

How EU MDR Changes Impact Supply Chain Agreements

The EU MDR supply chain agreement revolution has caught many medical device companies off guard. What started as a regulatory update has evolved into a comprehensive overhaul of how suppliers, distributors, and manufacturers must collaborate. If your contracts still look like they were written during the Medical Devices Directive era, you’re playing regulatory Russian roulette.

How EU MDR Changes Impact Supply Chain Agreements Read More »

Medical Device Distributor Responsibilities Under the EU MDR

Understand the responsibilities of medical device distributors under the EU MDR, including verification checks, storage and transport controls, traceability, complaint handling, non-compliant devices and cooperation with manufacturers, importers and competent authorities.

Medical Device Distributor Responsibilities Under the EU MDR Read More »

CHOOSING-THE-RIGHT-PARTNER

How to Select a Compliant EU Authorised Representative or Importer

Selecting an EU Authorised Representative or Importer is one of the most important regulatory decisions a non-EU medical device manufacturer will make. These organisations play a key role in market access, regulatory communications and ongoing compliance, so choosing the right partner requires more than simply comparing prices.

This guide explains how to assess prospective Authorised Representatives and Importers, the questions you should ask before appointment, the warning signs to avoid and the evidence you should review before signing an agreement.

How to Select a Compliant EU Authorised Representative or Importer Read More »

MDR/IVDR Compliance Obligations for Importers & Distributors

Understanding Your Obligations Under MDR/IVDR

Under the EU Medical Device Regulation (MDR) and In Vitro Diagnostic Regulation (IVDR), compliance isn’t just the manufacturer’s responsibility. Importers, distributors, and suppliers are now considered economic operators — and that means you are legally liable for specific obligations. From verifying CE marking and language requirements to registering devices in EUDAMED and maintaining long-term traceability, your role in the supply chain is critical.

Understanding Your Obligations Under MDR/IVDR Read More »

Illustration showing an EU MDR technical file being maintained throughout the medical device lifecycle, including document control, change management, clinical evaluation, risk management, post-market surveillance and ongoing regulatory compliance.

Building a Strong MDR Technical File: Maintaining Compliance Throughout the Medical Device Lifecycle

An MDR technical file is not a one-time submission created solely for CE marking. Under Regulation (EU) 2017/745, manufacturers must keep their technical documentation accurate, up to date and aligned with changes to the device, manufacturing processes, clinical evidence and post-market surveillance activities throughout the product lifecycle.

This guide explains how to maintain compliant technical documentation, when updates are required, and the practical steps manufacturers can take to ensure their technical file remains complete, consistent and inspection-ready.

Building a Strong MDR Technical File: Maintaining Compliance Throughout the Medical Device Lifecycle Read More »

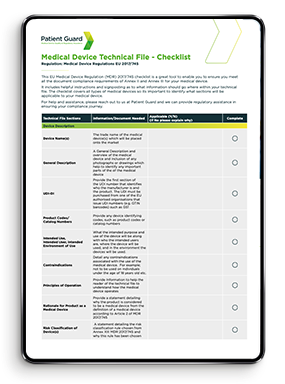
Get the Medical Device Technical Checklist

Thank you! The checklist is now ready to download.